The following publications have used the DeePMD-kit software. Publications that only mentioned the DeePMD-kit will not be included below.
We encourage explicitly mentioning DeePMD-kit with proper citations in your publications, so we can more easily find and list these publications.
Last update date: 06/11/2026
2026
Thermally and electronically triggered hydrogen shift within a CHCH2 species via surface-assisted hydrogen tunneling
Zhen Xu, Bin Jin, Zhen Zhu, Jinliang Pan, Zhichao Huang, Bin Di, Zhantao Peng, Wentao Li, Weijun Guo, Qiwei Chen, Yongfeng Wang, Pengju Ren, Xiaodong Wen, Yongwang Li, Shenzhen Xu, Xiong Zhou, Kai Wu
Nat. Commun., 2026, 17 (1), 9698.
DOI: 10.1038/s41467-026-77337-y
Light hydrogen isotopes in terrestrial core
Yu Zhang, Wenzhong Wang, Zhengbin Deng, Zhongqing Wu, Bowen Chen
Sci. Adv., 2026, 12 (28), eaec5685.
DOI: 10.1126/sciadv.aec5685
Thermal conductivity of seifertite and pyrite-type SiO 2 : A comparative study
Doyoon Park, Yihang Peng, Jie Deng
Phys. Rev. B, 2026, 114 (13), 134310.
DOI: 10.1103/gbbx-xkqh
A local structural basis to distinguish amorphous ices
Quinn M. Gallagher, Ryan J. Szukalo, Nicolas Giovambattista, Pablo G. Debenedetti, Michael A. Webb
Nat. Commun., 2026, 17 (1), 10754.
DOI: 10.1038/s41467-026-77279-5
Active Learning of a Neural Network Potential for Large-Scale Atomistic Simulations of Polymer Electrolyte Membranes
Yuta Yoshimoto, Naoki Matsumura, Meguru Yamazaki, Yuto Iwasaki, Hiroshi Nakao, Yasufumi Sakai
J. Phys. Chem. B, 2026, 130 (36), 9326–9337.
DOI: 10.1021/acs.jpcb.6c04744
Cross-scale simulation of electro-dewetting and interfacial behavior prediction based on microscopic diffusion coefficient transfer
Huang Peng, Ze-Jie Ke, Fei-Qiang Zhou, Jia-Yang Zhao, Rui Long, Yue Song
Acta Phys. Sin., 2026.
DOI: 10.7498/aps.76.20260763
A unified machine-learning framework for ab initio multiscale modeling of liquids
Anna T. Bui, Stephen J. Cox
Proc. National Acad. Sci. U. S. A., 2026, 123 (30), e2610049123.
DOI: 10.1073/pnas.2610049123
Disentangling cation-polyanion coupling reveals which anion motion dominates cation transport in solid electrolytes
Ke Li, Jitai Yang, Yu Zhai, Hui Li
Nat. Commun., 2026, 17 (1), 10455.
DOI: 10.1038/s41467-026-77273-x
Pressure-dependent melting and crystallization of B2-NiAl from neural- network molecular dynamics
A. S. Onegin, P. R. Levashov, N. M. Chtchelkatchev
J. Chem. Phys., 2026, 165 (12), 124513.
DOI: 10.1063/5.0347546
A Divide-and-Conquer Embedded Correlated Wavefunction Transfer Learning Approach for Chemically Accurate Condensed-Phase Interatomic Potentials: Liquid Water as a Test Case
Xuezhi Bian, Emily A. Carter
J. Phys. Chem. Lett., 2026, 17 (40), 11430–11442.
DOI: 10.1021/acs.jpclett.6c02897
Isotope-labeled growth histories reveal persistent chirality in individual carbon nanotubes despite catalyst evolution
Keigo Otsuka, Ryuji Fujiwara, Shigeo Maruyama
Nat. Commun., 2026, 17 (1), 8898.
DOI: 10.1038/s41467-026-77101-2
Stacking-engineered thermal transport and phonon filtering in rhenium disulfide
Yongjian Zhou, Haoran Cui, Zefang Ye, Jung-Fu Lin, Yan Wang, Yaguo Wang
Appl. Phys. Lett., 2026, 129 (8), 82107.
DOI: 10.1063/5.0342816
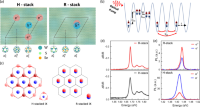
Robust valley-polarized excitonic Mott states and doublons enabled by stacking-controlled moir'e geometry
Hao-Tien Chu, Shou-Chien Chiu, Meng-Che Yeh, Yu-Wei Hsieh, Jia-Sian Su, Xiao-Wei Zhang, Jie-Yong Zeng, Po-Chun Huang, Si-Jie Chang, Kenji Watanabe, Takashi Taniguchi, Yunbo Ou, Seth Ariel Tongay, Ting Cao, Chaw-Keong Yong
Nat. Commun., 2026, 17 (1), 10865.
DOI: 10.1038/s41467-026-77907-0
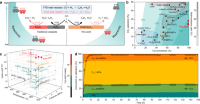
Constructing water-resistant phase-pure alloy carbide for boosting syngas to olefins
Ling Li, Heng Liang, Yuhan Peng, Linkai Wang, Ruoting Shan, Jida Wang, Xiang-Kui Gu, Yanfei Xu, Mingyue Ding
Nat. Commun., 2026, 17 (1), 9581.
DOI: 10.1038/s41467-026-76576-3
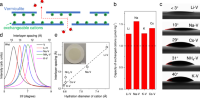
Ion-disrupted hydrogen-bond networks enable fast water transport under two-dimensional confinement
Mingyang Xia, Xianglong Du, Canbin Wang, Jinguo Liu, Zhihua Zhou, Ying-Ru Qiu, Yao-Hui Wang, Fujie Tang, Sheng Hu
Nat. Commun., 2026, 17 (1), 8585.
DOI: 10.1038/s41467-026-75604-6
Data-Efficient and Fast Machine Learning Molecular Dynamics through Integrated Active Learning and Knowledge Distillation
Xiliang Lian, Alfredo Pasquarello
J. Chem. Theory Comput., 2026, 22 (17), 8730–8739.
DOI: 10.1021/acs.jctc.6c00917
Deep-Learning Enabled Atomistic Understanding of Thermomechanical Behaviors and Fracture Mechanisms of High-Entropy Diboride (Hf0.2Zr0.2Ta0.2Ti0.2Nb0.2)B2
Xu Zhang, Bei Li, Meng Wang, Bo Liu, Ji Zou, Jianjun Li
Mater. (Basel Switz.), 2026, 19 (13), 2785.
DOI: 10.3390/ma19132785
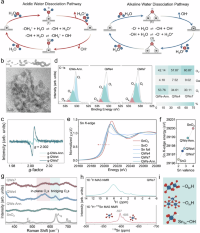
Bipolar membranes reveal surface-hydroxyl-structure-dependent water dissociation mechanism
Huijun Lv, Chao Yang, Minghao Sun, Yu Yang, Fengwang Li, Pengfei Ou, Yuhang Wang
Nat. Commun., 2026, 17 (1), 8284.
DOI: 10.1038/s41467-026-75170-x
Strong-bond networks and basal-cleavage resistance govern strength-toughness balance and elevated-temperature retention in Ta-based medium-entropy 211 MAX ceramics
Lu Liu, Qifeng Zhuang, Xiyu Xiao, Wei Ding, Haoran Zhou, Xuefang Yin, Kaicheng Zhang, Baoze Zhao, Shun Dong, Guobing Ying
J. Adv. Ceram., 2026, 15 (10), 9221368.
DOI: 10.26599/jac.2026.9221368
NNP/CG-MM: Embedding of All-Atom Neural Network Potentials into a Coarse-Grained Molecular Mechanics Environment
Kuntal Ghosh, Gregory A. Voth
J. Chem. Theory Comput., 2026, 22 (15), 7544–7557.
DOI: 10.1021/acs.jctc.6c00741
Modeling the Composition and Structure of Earth’s Inner Core Using Deep Machine Learning
V. E. Valiulin, A. V. Mikheyenkov, N. M. Chtchelkatchev, K. D. Litasov, V. V. Brazhkin
Jetp Lett., 2026, 124 (6), 427–433.
DOI: 10.1134/s0021364026607815
Multiscale Investigation of Triboelectric Nanogenerator: An Expanded DFT Calculation Based on Machine Learning
Yongsheng Huang, Mitsuhiro Matsumoto
Acs Omega, 2026, 11 (37), 56060–56070.
DOI: 10.1021/acsomega.6c06130
Turbulent-centrifugal-spun ceramic aerogels with elemental difference maximization design for thermal protection over 2000 K in air
Chuanyun Song, Yingde Zhao, Jiali Chen, Zhengli Yan, Shujin Laima, Geng Xue, Zelin Chou, Han Zhao, Yuanpeng Deng, Hongxuan Yu, Shixuan Dang, Duola Wang, Dizhou Liu, Cong Li, Jianing Zhang, Tiande Lin, Haiyang Zhao, Shaoyi Zhang, Jingran Guo, Hui Li, Xiang Xu
Sci. Adv., 2026, 12 (30), eaee9953.
DOI: 10.1126/sciadv.aee9953
Machine Learning Potential for Calcium Silicate Hydrates With Broad Compositional and Structural Diversity
Yunjian Li, Cheng Chen, Roland J-M Pellenq, Zongjin Li, Jiaping Liu
Adv. Sci. (Weinheim Baden-Wurtt.- Ger.), 2026, e77942.
DOI: 10.1002/advs.77942
Tracing the Staging Dynamics and Li+ Kinetics in Graphite Anodes via Automated Continuous (De)Lithiation Simulation
Liqi Wang, Xuhe Gong, Zicun Li, Ruijuan Xiao, Hong Li
Acs Appl. Energy Mater., 2026, 9 (17), 11989–12010.
DOI: 10.1021/acsaem.6c02065
Interface-dependent phase transitions and ultrafast hydrogen superionic diffusion of H 2 O ice
Pengfei Hou, Yumiao Tian, Zifeng Liu, Junwen Duan, Hanyu Liu, Xing Meng, Russell J. Hemley, Yanming Ma
Phys. Rev. B, 2026, 113 (18), 184103.
DOI: 10.1103/h61k-f3mj
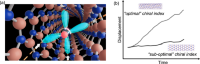
Direction-specific enhanced diffusion of CO2 in chiral hexagonal boron nitride nanotubes
Manh-Thuong Nguyen, David J. Heldebrant, Jian Liu, Abhoyjit S. Bhown, Zhijie Xu
Nat. Commun., 2026, 17 (1), 4771.
DOI: 10.1038/s41467-026-72123-2
Iron-Catalyzed Chlorination of Titanium Oxides in Molten Salts: A Deep Neural Network-Based Mechanistic Study
Liangliang Gu, Jie Zhou, Wei Liu, Yuanyuan Chen, Linfei Li, Ronggang Sun, Rong Yu, Xiumin Chen, Yunmin Chen
Mater. (Basel Switz.), 2026, 19 (9), 1746.
DOI: 10.3390/ma19091746
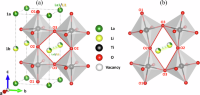
Engineering ionic pathways by phonon-structure connection in perovskite solid electrolyte Li0.29La0.57TiO3
Samiran Malgope, Mayanak K. Gupta, Ranjan Mittal, Olivier Delaire, Rakesh Shukla, Srungarpu N. Achary, Mohamed Aouane, Avesh K. Tyagi, Samrath L. Chaplot
Commun. Mater., 2026, 7 (1), 165.
DOI: 10.1038/s43246-026-01164-3
SPARC: An Automated Workflow Toolkit for Accelerated Active Learning of Reactive Machine Learning Interatomic Potentials
Rahul Verma, Nisarg Joshi, Jim Pfaendtner
Joss, 2026, 11 (121), 9468.
DOI: 10.21105/joss.09468
A Deep Neural Network Potential Study on the Mechanical and Thermophysical Properties of β-HMX Molecular Crystal
Jiyuan Wei, Yifeng Dong, Teng Zhang, Hao Liu, Yao Long, Ying Li, Jun Chen
Acs Omega, 2026, 11 (20), 29733–29747.
DOI: 10.1021/acsomega.5c13571
Precessional Dynamics of Octahedra in CsPbBr3
Lucas Martin Farigliano, Marcio S Gomes-Filho, Alexandre Reily Rocha, Gustavo Martini Dalpian
J. Chem. Theory Comput., 2026, 22 (8), 4125–4135.
DOI: 10.1021/acs.jctc.5c02166
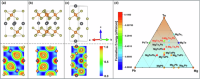
Ductile Mg-Te-Pb thermoelectric materials with ultralow lattice thermal conductivity predicted by a deep learning potential model
Xin-Xuan Wang, Zhen-Shuai Lei, Wen-Juan Li, Xiao-Bin Feng, Gang Chen, Peng-Cheng Zhai, Bo Duan, Guo-Dong Li, Qing-Jie Zhang
npj Comput. Mater., 2026, 12 (1), 270.
DOI: 10.1038/s41524-026-02149-z
Ion Distribution and Cation Exchange at Mica-Electrolyte Interfaces Probed with Deep Potential Molecular Dynamics
Sanghyun J. Park, Abhinav S. Raman, Annabella Selloni
Chem. Mater., 2026, 38 (10), 5049–5057.
DOI: 10.1021/acs.chemmater.6c00214
Understanding the density maximum of water with machine-learned potentials
Yizhi Song, Renxi Liu, Chunyi Zhang, Yifan Li, Biswajit Santra, Mohan Chen, Michael L. Klein, Xifan Wu
Sci. Adv., 2026, 12 (22), eaec6748.
DOI: 10.1126/sciadv.aec6748
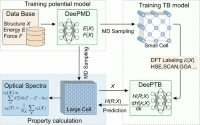
Deep neural network for phonon-assisted optical spectra of semiconductors at finite temperatures
Shengguo Yang, Shishir Kumar Pandey, Zhanghao Zhouyin, Qiangqiang Gu
npj Comput. Mater., 2026, 12 (1), 268.
DOI: 10.1038/s41524-026-02137-3
Data-Driven Construction of Machine-Learning-Based Interatomic Potentials for Gas-Surface Scattering Dynamics: The Case of NO on Graphite
Samuel Del Fr'e, Gilberto A. Alou Angulo, Maurice Monnerville, Alejandro Rivero Santamar'\ia
J. Phys. Chem. C, 2026, 130 (20), 7037–7050.
DOI: 10.1021/acs.jpcc.6c01815
Interlayer coupling driven stabilization and superconductivity in bilayer CoTe 2
Wenping Chen, Ziyun Zhang, Feipeng Zheng
Phys. Rev. B, 2026, 114 (5), 55413.
DOI: 10.1103/l89c-t2s4
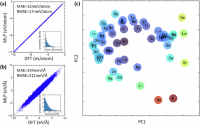
A domain-specific machine learning potential model for metallic materials spanning 53 elements
Xin-Yang Li, Jing Li, Yi-Nan Wang, Na-Min Xiao, Wen-Yue Zhao, Zhang-Zhi Shi, Xin-Fu Gu, Fu-Zhi Dai, Lu-Ning Wang
npj Comput. Mater., 2026, 12 (1), 208.
DOI: 10.1038/s41524-026-02072-3
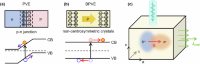
Accelerating discovery of infrared nonlinear optical materials with large shift current via high-throughput screening
Aiqin Yang, Dian Jin, Mingkang Liu, Daye Zheng, Qi Wang, Qiangqiang Gu, Jian-Hua Jiang
npj Comput. Mater., 2026, 12 (1), 188.
DOI: 10.1038/s41524-026-02064-3
Ionic interdiffusion at cathode|solid-electrolyte interface: A machine learning-assisted multiscale investigation and mitigation strategies
Musawenkosi K. Ncube, Pallab Barai, Selva Chandrasekaran Selvaraj, Larry A. Curtiss, Anh T. Ngo, Venkat Srinivasan
J. Energy Storage, 2026, 169, 122678.
DOI: 10.1016/j.est.2026.122678
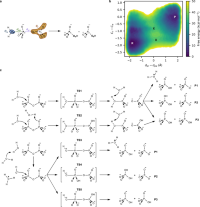
Protonation and magnesium ions shape the transition state diversity of phosphoanhydride hydrolysis in water
Adri'an Garc'\ia-Mart'\inez, Miguel de la Puente, Marion Cunin, Kirill Zinovjev, J. Javier Ruiz-Pern'\ia, Damien Laage, I\~naki Tu\~n'on
Nat. Commun., 2026, 17 (1), 6878.
DOI: 10.1038/s41467-026-73697-7
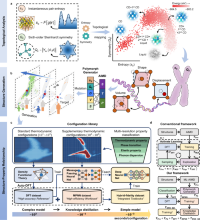
Navigating polymorph generation and distilled-potential development via entropy-symmetry landscapes for metal plasticity mechanisms
Zeyuan Li, Taiqiao Liu, Xuhao Wan, Songpeng Zhao, Zhaofu Zhang, E. Zhou, Wei Yu, Yijing Zuo, John Robertson, Sheng Liu, Yuzheng Guo
Nat. Commun., 2026, 17 (1), 5070.
DOI: 10.1038/s41467-026-73188-9
Automating Computational Chemistry Workflows via OpenClaw and Domain- Specific Skills
Mingwei Ding, Chen Huang, Yibo Hu, Yifan Li, Zitian Lu, Xingtai Yu, Duo Zhang, Wenxi Zhai, Tong Zhu, Qiangqiang Gu, Jinzhe Zeng
J. Chem. Theory Comput., 2026, 22 (12), 5919–5929.
DOI: 10.1021/acs.jctc.6c00622
Rejuvenation of Mechanical Fatigue Resistance in 2D Ferroelectric CuInP2S6 by Reversing Ionic Motion
Xiqi Wu, Shuai Han, Zhaoheng Zhang, Yafei Wang, Jiahao Li, Zhao Zhang, Dong Wang, Xinshuai Peng, Yixuan She, Xiang Gao, Chongjian Zhou, Guorui Wang, Zhong Zhang
Small (Weinheim Bergstr. Ger.), 2026, 22 (30), e73338.
DOI: 10.1002/smll.73338
Enhanced graphene-water thermal transport via edge functionalization without compromising in-plane thermal conductivity
John Crosby, Haoran Cui, Mehrab Lotfpour, Yan Wang, Lei Cao
2D Mater., 2026, 13 (3), 35005.
DOI: 10.1088/2053-1583/ae780d
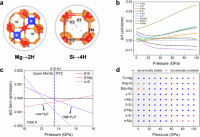
Hydrogen site-dependent physical properties of hydrous magnesium silicates in the mantle transition zone
Zifan Wang, Yu He, Ho-Kwang Mao, Duck Young Kim
Nat. Commun., 2026, 17 (1), 5970.
DOI: 10.1038/s41467-026-72807-9
Transfer Learning Meets Embedded Correlated Wavefunction Theory for Chemically Accurate Molecular Simulations: Application to Calcium Carbonate Ion Pairing
Xuezhi Bian, Emily A. Carter
J. Chem. Theory Comput., 2026, 22 (10), 5174–5184.
DOI: 10.1021/acs.jctc.6c00403
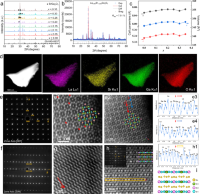
Interstitial cation defect chemistry and correlated disorder in melilite solid electrolytes
Xiao Ma, Xiaohui Li, Xianyi Wei, Cheng Li, C'ecile Genevois, Mathieu Allix, Xiaoge Wang, Qilong Gao, Xiaoming Wang, Sihao Deng, Lunhua He, Lu Liang, Qiang Li, Xianran Xing, Xiaojun Kuang
Nat. Commun., 2026, 17 (1), 5722.
DOI: 10.1038/s41467-026-72322-x
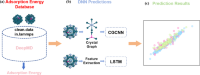
Fast and accurate prediction of adsorption energy of AgPd nanoalloys by deep learning potentials and neural networks
Wanxuan Zhang, Shuang Shan, Fanzhe Ma, Xiaoqing Liu, Peng Xu, Fuyi Chen
Sci. Rep., 2026, 16 (1), 652.
DOI: 10.1038/s41598-025-05309-1
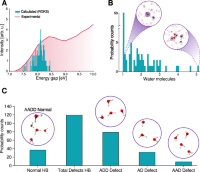
Simulating the Photochemical Birth of the Hydrated Electron in Liquid Water
Gonzalo D'\iaz Mir'on, Cesare Malosso, Solana Di Pino, Colin K. Egan, Diganta Dasgupta, Christopher J. Mundy, Ali Hassanali
Nat. Commun., 2026, 17 (1), 3764.
DOI: 10.1038/s41467-026-70045-7
On the boroxol ring fraction in melt-quenched B2O3 glass: Insights from machine learning potentials
Debendra Meher, Nikhil V. S. Avula, Sundaram Balasubramanian
J. Chem. Phys., 2026, 164 (12), 124504.
DOI: 10.1063/5.0321441
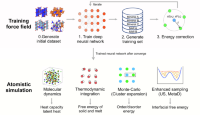
A general purposed machine learning interatomic potential for Mg-Al- Si-O system suitable for Earth materials at high pressure and temperature conditions
Xin Zhong, Yifan Li, Timm John
npj Comput. Mater., 2026, 12 (1), 141.
DOI: 10.1038/s41524-026-02056-3
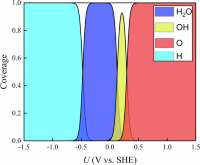
Microkinetic modeling of acidic corrosion from first principles and machine-learning molecular dynamics
Ergen Bao, Wenjing Xu, Hui Ma, Yueqi Si, Ijaz Shahid, Yutong Huo, Peitao Liu, Yan Sun, Xing-Qiu Chen
npj Comput. Mater., 2026, 12 (1), 185.
DOI: 10.1038/s41524-026-02047-4
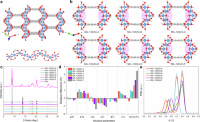
Decoding local framework dynamics in the ultra-small pore MOF MIL-120(Al) CO2 adsorbent using machine-learning potential
Dong Fan, Felipe Lopes Oliveira, Satyanarayana Bonakala, Mohammad Wahiduzzaman, Guillaume Maurin
Nat. Commun., 2026, 17 (1), 3235.
DOI: 10.1038/s41467-026-69993-x
Absence of dehydration due to superionic transition at Earth's core- mantle boundary
Yu He, Wei Zhang, Qingyang Hu, Shichuan Sun, Jiaqi Hu, Daohong Liu, Li Zhou, Lidong Dai, Duck Young Kim, Simon A. T. Redfern, Yun Liu, Heping Li, Ho-Kwang Mao
Sci. Adv., 2026, 12 (5), eaeb3006.
DOI: 10.1126/sciadv.aeb3006
Dynamic Potassium Segregation Drives Cu2+ \textrightarrow Cu+ Transition and Redefines Alkali-Metal Promotion in Selective Propylene Epoxidation
Dingming Chen, Danfeng Xiong, Haifeng Wang
JACS Au, 2026, 6 (4), 2580–2590.
DOI: 10.1021/jacsau.6c00225
Spectral Similarity Masks Structural Diversity at Hydrophobic Water Interfaces
Yong Wang, Yifan Li, Linhan Du, Chunyi Zhang, Lorenzo Agosta, Marcos Calegari Andrade, Annabella Selloni, Roberto Car
Phys. Rev. Lett., 2026, 136 (5), 56202.
DOI: 10.1103/w27h-sclq
Machine learning potential as a guide for eutectic in ultra-refractory multicomponent ceramics
V. E. Valiulin, A. V. Mikheyenkov, N. M. Chtchelkatchev, E. A. Levashov
J. Chem. Phys., 2026, 164 (5), 54504.
DOI: 10.1063/5.0309969
Ground state of CuInP 2 S 6 thin films: A study of the deep potential method
Shengxian Li, Jiaren Yuan, Tao Ouyang, Anlian Pan, Mingxing Chen
Phys. Rev. B, 2026, 113 (12), 125417.
DOI: 10.1103/rp2y-2b5j
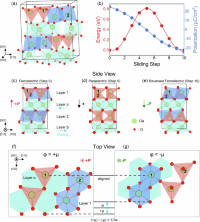
Origin of suppressed ferroelectricity in κ-Ga2O3: interplay between polarization and lattice domain walls
Yonghao Zhu, Wen-Hao Liu, Run Long, Lin-Wang Wang, Jun-Wei Luo, Zhi Wang
npj Comput. Mater., 2026, 12 (1), 155.
DOI: 10.1038/s41524-026-02022-z
Tuning Molecular Thermal Conductance through End-Group Modification and Halogen Substitution
Jonathan J. Wang, Dvira Segal
J. Phys. Chem. C, 2026, 130 (12), 4618–4626.
DOI: 10.1021/acs.jpcc.5c08503

Dynamic metal-ligand coordination in solution: a ligand-exchange relay mechanism for gold(i) sulfido complex transformation
Yining Zhang, Guoao Li, Manyi Yang, Shuhua Li
Chem. Sci., 2026.
DOI: 10.1039/d6sc06455h
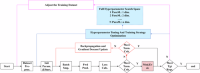
Short bond evaluation method for rapidly assessing the generalization ability of deep neural network potential function models and its effectiveness verification
Xiumin Chen, Yunmin Chen, Jie Zhou
npj Comput. Mater., 2026, 12 (1), 90.
DOI: 10.1038/s41524-026-01957-7

Simulating enzyme catalysis with electrostatically embedded machine learning potentials
Valentin Gradisteanu, Elliot W. Chan, Lester Hedges, Meritxell Malagarriga, Rolf David, Miguel de la Puente, Damien Laage, I\~naki Tu\~n'on, Marc W. van der Kamp, Kirill Zinovjev
Chem. Sci., 2026, 17 (17), 8542–8556.
DOI: 10.1039/d6sc01156j
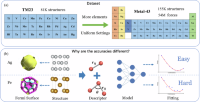
Origin of the machine learning forces field errors across metal elements
Xingze Geng, Wentao Zhang, Lin-Wang Wang, Xiangying Meng
npj Comput. Mater., 2026, 12 (1), 102.
DOI: 10.1038/s41524-026-01977-3

Solvent-temperature coupled hydride transfer controls product selectivity in the HSiR3/KO t Bu catalytic system
Haojiang Yao, Zhenhao Zhou, Ye Liu, Dong H. Zhang, Guohui Li
Chem. Sci., 2026, 17 (20), 10092–10103.
DOI: 10.1039/d6sc00107f
Deep learning potential for accurate shock response simulations in tin
Yixin Chen, Xiaoyang Wang, Wanghui Li, Mohan Chen, Han Wang
Phys. Rev. Mater., 2026, 10 (2), 23605.
DOI: 10.1103/6psb-3ksf
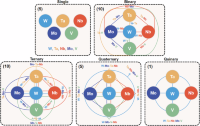
Scaling reliable interatomic potentials to complex nuclear alloys via pretrained atomic models
Mingxuan Jiang, Biao Xu, Yixin Deng, Shihua Ma, Ji-Jung Kai, Fei Gao, Huiqiu Deng
npj Comput. Mater., 2026, 12 (1), 82.
DOI: 10.1038/s41524-025-01950-6
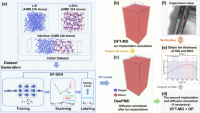
Deep learning modeling of oxygen redistribution and thermal transport in silicon on insulator and buried oxide layers
Xiuwen Yan, Mingzhu Liu, Wenjin Cheng, Yong Sun, Yiwen Su, Xinzhong Wang, Zhuo Tang, Libo Peng, Guangping Zheng, Jiashu Chen
Commun. Mater., 2026, 7 (1), 80.
DOI: 10.1038/s43246-026-01130-z
Computational discovery of high-temperature superconducting ternary hydrides via deep learning
Xiaoyang Wang, Chengqian Zhang, Zhenyu Wang, Hanyu Liu, Jian Lv, Han Wang, E. Weinan, Yanming Ma
National Sci. Rev., 2026, 13 (6), nwag030.
DOI: 10.1093/nsr/nwag030
Beyond geometry orders: uncovering bonding-heterogeneity-dominated structure-relaxation coupling in glasses
Liang Gao, Jia-Qi Gao, Qing-Zhou Bu, Qun Yang, Yang Sun, Kai-Ming Ho, Qi Wang, Jeppe C. Dyre, Hai-Bin Yu
National Sci. Rev., 2026, 13 (5), nwag006.
DOI: 10.1093/nsr/nwag006
Mechanistic Analysis of Large Atomic Models of Molten Salt
Yuliang Guo, Xiaobo Sun, Xiaoli Xi, Zuoren Nie
Adv. Sci. (Weinheim Baden-Wurtt.- Ger.), 2026, 13 (22), e22313.
DOI: 10.1002/advs.202522313
Energy landscape statistics and thermodynamics of a machine-learned model of water
Ryan J. Szukalo, Andreas Neophytou, Axel Gomez, Nicolas Giovambattista, Francesco Sciortino, Pablo G. Debenedetti
Proc. National Acad. Sci. U. S. A., 2026, 123 (13), e2534303123.
DOI: 10.1073/pnas.2534303123
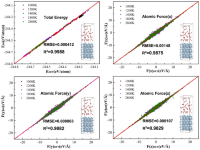
Efficient removal of AlN from secondary aluminum dross using binary fluoride: a theoretical and experimental study
Tianshuang Li, Zhaohui Guo, Hong Qin, Huimin Xie
Sci. Rep., 2026, 16 (1), 12986.
DOI: 10.1038/s41598-026-43443-6
Machine Learning Discovery of Record-Low Lattice Thermal Conductivity in Double Perovskites
Md Zaibul Anam, Alejandro Rodriguez, Riccardo Rurali, Ming Hu
Adv. Sci. (Weinheim Baden-Wurtt.- Ger.), 2026, 13 (18), e15766.
DOI: 10.1002/advs.202515766
2025
Uncovering coupled ionic-polaronic dynamics and interfacial enhancement in Li x FePO 4
Fengyu Xie, Yuxiang Gao, Ruoyu Wang, Zhicheng Zhong
Phys. Rev. B, 2026, 113 (2), 24105.
DOI: 10.1103/rnrn-zfqq
Revealing the dynamic responses of Pb under shock loading based on DFT-accuracy machine learning potential
Enze En Ze Hou Hou, Xiaoyang Xiao Yang Wang Wang, Han Han Wang Wang
Chin. Phys. B, 2026, 35 (1), 18701.
DOI: 10.1088/1674-1056/ae1726
Interlayer Electron Redistribution Engineering for Ultralow Friction in 2D Electrides
Jingcheng Qi, Giuliana Materzanini, Gian-Marco Rignanese, Maria Clelia Righi, Junjie Wang
Adv. Funct. Mater., 2026, 36 (26), e25865.
DOI: 10.1002/adfm.202525865
Accuracy and limitations of machine-learned interatomic potentials for magnetic systems: A case study on Fe-Cr-C
E. O. Khazieva, N. M. Chtchelkatchev, N. N. Katkov, R. E. Ryltsev
Phys. Rev. E, 2025, 112 (5-2), 55302.
DOI: 10.1103/913y-p6qf
Influence of Aluminum Distribution in Cu-MOR Systems on Methane-to- Methanol Conversion: A Combined Experimental and Theoretical Study
Peter N. Njoroge, Bj\orn Gading Solemsli, Asanka Wijerathne, Izar Capel Berdiell, Agnieszka Seremak, Mario Chiesa, Yu-Kai Liao, Beatrice Garetto, Nishant Patel, Karoline Kvande, Elisa Borfecchia, Christopher Paolucci, Unni Olsbye, Pablo Beato, Stian Svelle, Sebastian Prodinger
J. Phys. Chem. C, 2025, 129 (44), 20044–20056.
DOI: 10.1021/acs.jpcc.5c06045
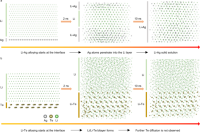
A porous tellurium interlayer for high-power and long-cycling garnet- based quasi-solid-state lithium-metal batteries
Ju-Sik Kim, Gabin Yoon, Yong Su Kim, Tae-Hee Kim, Sewon Kim, Jirae Kim, Junhyeong Lee, Ryounghee Kim, Myung-Jin Lee, Nobuyoshi Yashiro, Shinya Suzuki, Tetsuya Asano, Michael Badding, Zhen Song, Sung Heo
Nat. Commun., 2025, 16 (1), 11471.
DOI: 10.1038/s41467-025-66308-4
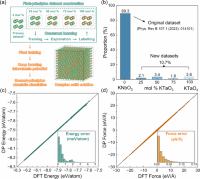
Machine learning-enabled atomistic insights into phase boundary engineering of solid-solution ferroelectrics
Weiru Wen, Fanda Zeng, Ben Xu, Ke Bi, Zhipeng Xing, Hao-Cheng Thong, Ke Wang
npj Comput. Mater., 2025, 11 (1), 343.
DOI: 10.1038/s41524-025-01829-6
Simulations of dielectric permittivity of water by machine learned potentials with long-range Coulombic interactions
Kehan Cai, Chunyi Zhang, Xifan Wu
J. Chem. Phys., 2025, 163 (18), 184503.
DOI: 10.1063/5.0294554
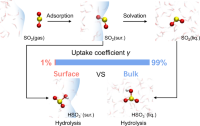
Bulk Phase Dominates Sulfur Dioxide Hydrolysis over Interfacial Processes
Mile Du, Manyi Yang, Han Wang, Yu Song, Tong Zhu
Nat. Commun., 2025, 17 (1), 558.
DOI: 10.1038/s41467-025-67250-1

Twist-angle transferable continuum model and second flat Chern band in twisted MoTe2 and WSe2
Xiao-Wei Zhang, Kaijie Yang, Chong Wang, Xiaoyu Liu, Ting Cao, Di Xiao
npj Quantum Mater., 2025, 10 (1), 110.
DOI: 10.1038/s41535-025-00828-6
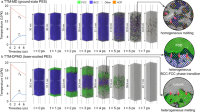
Picosecond-scale heterogeneous melting of metals at extreme non- equilibrium states
Qiyu Zeng, Xiaoxiang Yu, Bo Chen, Shen Zhang, Kaiguo Chen, Dongdong Kang, Jiayu Dai
Nat. Commun., 2025, 16 (1), 10464.
DOI: 10.1038/s41467-025-65485-6
Pre-training, fine-tuning, and distillation (PFD): Automatically generating machine learning force fields from universal models
Ruoyu Wang, Yuxiang Gao, Hongyu Wu, Zhicheng Zhong
Phys. Rev. Mater., 2025, 9 (11), 113802.
DOI: 10.1103/sbz6-btz8
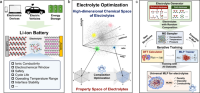
Domain oriented universal machine learning potential enables fast exploration of chemical space of battery electrolytes
Feng Wang, Yu-Hang Tang, Ze-Bing Ma, Yu-Cheng Jin, Jun Cheng
Nat. Commun., 2025, 17 (1), 1226.
DOI: 10.1038/s41467-025-67982-0
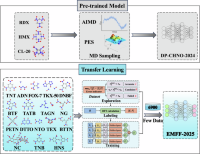
EMFF-2025: a general neural network potential for energetic materials with C, H, N, and O elements
Mingjie Wen, Jiahe Han, Wenjuan Li, Xiaoya Chang, Qingzhao Chu, Dongping Chen
npj Comput. Mater., 2025, 11 (1), 333.
DOI: 10.1038/s41524-025-01809-w
Pressure-driven electronegativity inversion in alkali liquids
Chenxu Han, Hongxiang Zong, Xiangdong Ding, Jun Sun, Graeme J. Ackland
Proc. National Acad. Sci. U. S. A., 2025, 122 (26), e2424701122.
DOI: 10.1073/pnas.2424701122
Harnessing artificial intelligence for modeling amorphous and amorphous porous palladium: A deep neural network approach
Isa'\ias Rodr'\iguez
Mrs Commun., 2025, 15 (4), 682–689.
DOI: 10.1557/s43579-025-00738-5
Ferroelectric Domains and Evolution Dynamics in Twisted CuInP2S6 Bilayers
Dongyu Bai, Junxian Liu, Yihan Nie, Yuantong Gu, Dongchen Qi, Arkady V. Krasheninnikov, Liangzhi Kou
Small Methods, 2026, 10 (1), e2500683.
DOI: 10.1002/smtd.202500683
Transferability of the chemical-bond-based machine learning model for dipole moment: The GHz to THz dielectric properties of liquid propylene glycol and polypropylene glycol
Tomohito Amano, Tamio Yamazaki, Naoki Matsumura, Yuta Yoshimoto, Shinji Tsuneyuki
Phys. Rev. B, 2025, 111 (16), 165149.
DOI: 10.1103/physrevb.111.165149
Superionicity of oxygen-deficient davemaoite and its impact on the deep-Earth oxidation cycle
Zifan Wang, Yu He, Ho-Kwang Mao, Duck Young Kim
Sci. Adv., 2025, 11 (22), eadu8401.
DOI: 10.1126/sciadv.adu8401
Charge orders in fully intercalated bilayer TaSe 2 : Dependence on interlayer stacking and intercalation sites
Yuhui Yan, Lingxiao Xiong, Feipeng Zheng
Phys. Rev. B, 2025, 111 (20), 205437.
DOI: 10.1103/physrevb.111.205437
Accurate Deep Potential Model of Temperature-Dependent Elastic Constants for Phosphorus-Doped Silicon
Miao Gao, Xiaorui Bie, Yi Wang, Yuhang Li, Zhaoyang Zhai, Haoqi Lyu, Xudong Zou
Nanomater. (Basel Switz.), 2025, 15 (10), 769.
DOI: 10.3390/nano15100769
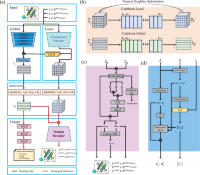
Massive discovery of crystal structures across dimensionalities by leveraging vector quantization
ZiJie Qiu, Luozhijie Jin, Zijian Du, Hongyu Chen, Guanyao Mao, Yan Cen, Siqi Sun, Yongfeng Mei, Hao Zhang
npj Comput. Mater., 2025, 11 (1), 184.
DOI: 10.1038/s41524-025-01613-6
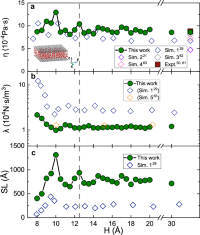
Synergistic effects of confinement size and interface on anomalous ultrafast transport in nanofluidics
Weiwen Pu, Zhaoru Sun
Commun. Phys., 2025, 8 (1), 167.
DOI: 10.1038/s42005-025-02085-2
Observing nucleation and crystallization of rock salt LiF from molten state through molecular dynamics simulations with refined machine- learned force field
Boyuan Xu, Liyi Bai, Shenzhen Xu, Qisheng Wu
J. Chem. Phys., 2025, 162 (23), 234705.
DOI: 10.1063/5.0276535
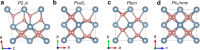
Unexpected density functional dependence of the antipolar \$\$\textbackslashboldsymbol{Pbcn}\$\$ phase in HfO2
Di Fan, Tianyuan Zhu, Shi Liu
npj Comput. Mater., 2025, 11 (1), 140.
DOI: 10.1038/s41524-025-01647-w
Neural Canonical Transformations for Quantum Anharmonic Solids of Lithium
Qi Zhang, Xiaoyang Wang, Rong Shi, Xinguo Ren, Han Wang, Lei Wang
Phys. Rev. Lett., 2025, 134 (24), 246101.
DOI: 10.1103/p3th-25bc
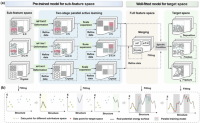
Transferable machine learning model for multi-target nanoscale simulations in hydrogen-carbon system from crystal to amorphous
Weiqi Chen, Zhiyue Xu, Kang Wang, Lei Gao, Aisheng Song, Tianbao Ma
npj Comput. Mater., 2025, 11 (1), 119.
DOI: 10.1038/s41524-025-01629-y
Exploring homogeneous ice nucleation through molecular dynamics simulations with ab initio accuracy
Mingyi Chen, Lin Tan, Han Wang, Linfeng Zhang, Haiyang Niu
Phys. Rev. B, 2025, 111 (13), 134116.
DOI: 10.1103/physrevb.111.134116
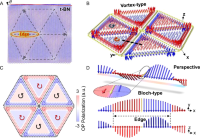
Edge polarization topology integrated with sliding ferroelectricity in Moir'e system
Wen-Cheng Fan, Zhao Guan, Lu-Qi Wei, Hao-Wen Xu, Wen-Yi Tong, Ming Tian, Neng Wan, Cheng-Shi Yao, Jun-Ding Zheng, Bin-Bin Chen, Ping-Hua Xiang, Ni Zhong, Chun-Gang Duan
Nat. Commun., 2025, 16 (1), 3557.
DOI: 10.1038/s41467-025-58877-1
Generator of Neural Network Potential for Molecular Dynamics: Constructing Robust and Accurate Potentials with Active Learning for Nanosecond-Scale Simulations
Naoki Matsumura, Yuta Yoshimoto, Tamio Yamazaki, Tomohito Amano, Tomoyuki Noda, Naoki Ebata, Takatoshi Kasano, Yasufumi Sakai
J. Chem. Theory Comput., 2025, 21 (8), 3832–3846.
DOI: 10.1021/acs.jctc.4c01613
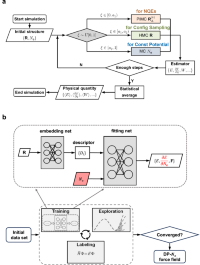
Probing nuclear quantum effects in electrocatalysis via a machine-learning enhanced grand canonical constant potential approach
Menglin Sun, Bin Jin, Xiaolong Yang, Shenzhen Xu
Nature Communications, 2025, 16, 3600.
DOI: 10.1038/s41467-025-58871-7
Machine Learning Potential for Electrochemical Interfaces with Hybrid Representation of Dielectric Response
Jia-Xin Zhu, Jun Cheng
Phys. Rev. Lett., 2025, 135 (1), 18003.
DOI: 10.1103/48ct-3jxm
Application of Deep-Learning Potential in Simulating the Structural and Physical Characteristics of Platinum
Keyuan Chen, Xingkao Zhang, Li Ma, Jueyi Ye, Qi Qiu, Haoxiang Zhang, Ju Rong, Yudong Sui, Xiaohua Yu, Jing Feng
Cmc, 2025, 83 (1), 685–700.
DOI: 10.32604/cmc.2025.060713
An interpretable DeePMD-kit performance model for emerging supercomputers
Xiangyu Meng, Xun Wang, Mingzhen Li, Guangming Tan, Weile Jia
Ccf Trans. Hpc, 2025, 7 (2), 155–168.
DOI: 10.1007/s42514-024-00209-8
Liquid-Vapor Phase Equilibrium in Molten Aluminum Chloride (AlCl3) Enabled by Machine Learning Interatomic Potentials
Rajni Chahal, Luke D. Gibson, Santanu Roy, Vyacheslav S. Bryantsev
J. Phys. Chem. B, 2025, 129 (3), 952–964.
DOI: 10.1021/acs.jpcb.4c06450
Primordial hydrogen partitioning at Earth’s core–mantle boundary: Multicomponent effects revealed by machine learning-augmented first-principles simulations
ZePing Jiang, Key Laboratory of Planetary Science, Institute of Geology Frontier Technology, China Beijing 100029 Chinese Academy of Sciences Geophysics, YuYang He, ZhiGang Zhang, College of Earth, China Beijing 100029 University of Chinese Academy of Sciences Planetary Sciences
Earth Planet. Phys., 2025, 9 (5), 1001–1009.
DOI: 10.26464/epp2025055
Structural and mechanical properties of W-Cu compounds characterized by a neural-network-based potential
Jianchuan Liu, Tao Chen, Sheng Mao, Mohan Chen
Comput. Mater. Sci., 2025, 253, 113825.
DOI: 10.1016/j.commatsci.2025.113825
Edge perfection of two-dimensional materials
Pengjie Shi, Zhiping Xu
Mrs Bull., 2025, 50 (5), 549–556.
DOI: 10.1557/s43577-024-00855-x
Pressure-induced structural and dielectric changes in liquid water at room temperature
Yizhi Song, Xifan Wu
J. Chem. Phys., 2025, 162 (11), 114508.
DOI: 10.1063/5.0247114
Slowly quenched, high pressure glassy B2O3 at DFT accuracy
Debendra Meher, Nikhil V. S. Avula, Sundaram Balasubramanian
J. Chem. Phys., 2025, 162 (4), 44503.
DOI: 10.1063/5.0240030

Grain boundary diffusion cannot explain the W isotope heterogeneities of the deep mantle
Yihang Peng, Takashi Yoshino, Jie Deng
Nat. Commun., 2025, 16 (1), 1866.
DOI: 10.1038/s41467-025-57120-1
Glassy dynamics in a glass-forming liquid: A first-principles study of toluene
Florian Pabst, Stefano Baroni
Phys. Rev. E, 2025, 111 (2), L023401.
DOI: 10.1103/physreve.111.l023401
Mechanistic insights into temperature effects for ionic conductivity in Li6PS5Cl
Zicun Li, Jianxing Huang, Xinguo Ren, Jinbin Li, Ruijuan Xiao, Hong Li
Journal of Power Sources, 2025, 640, 236632.
DOI: 10.1016/j.jpowsour.2025.236632
Molecular Fingerprints of Ice Surfaces in Sum Frequency Generation Spectra: A First-Principles Machine Learning Study
Margaret L. Berrens, Marcos F. Calegari Andrade, John T. Fourkas, Tuan Anh Pham, Davide Donadio
JACS Au, 2025, 5 (3), 1173–1183.
DOI: 10.1021/jacsau.4c00957

Insights into defect cluster formation in non-stoichiometric wustite (Fe1$-$xO) at elevated temperatures: accurate force field from deep learning
Zeng Liang, Kejiang Li, Jianliang Zhang, Alberto N. Conejo
npj Comput. Mater., 2025, 11 (1), 32.
DOI: 10.1038/s41524-025-01527-3
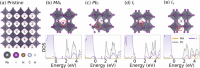
Sub-bandgap charge harvesting and energy up-conversion in metal halide perovskites: ab initio quantum dynamics
Bipeng Wang, Weibin Chu, Yifan Wu, Wissam A. Saidi, Oleg V. Prezhdo
npj Comput. Mater., 2025, 11 (1), 11.
DOI: 10.1038/s41524-024-01467-4
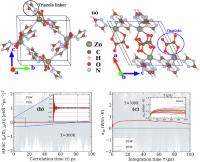
Prediction of thermal conductivity in CALF-20 with first-principles accuracy via machine learning interatomic potentials
Soham Mandal, Prabal K. Maiti
Commun. Mater., 2025, 6 (1), 22.
DOI: 10.1038/s43246-025-00745-y
Propensity of Water Self-Ions at Air(Oil)-Water Interfaces Revealed by Deep Potential Molecular Dynamics with Enhanced Sampling
Pengchao Zhang, Xuefei Xu
Langmuir: Acs J. Surf. Colloids, 2025, 41 (5), 3675–3683.
DOI: 10.1021/acs.langmuir.4c05004
Chirality-Dependent Kinetics of Single-Walled Carbon Nanotubes from Machine-Learning Force Fields
Sida Sun, Shigeo Maruyama, Yan Li
J. Am. Chem. Soc., 2025, 147 (8), 7103–7112.
DOI: 10.1021/jacs.4c18769
Switching Two-Dimensional Sliding Ferroelectrics by Mechanical Bending
Ri He, Hua Wang, Fenglin Deng, Yuxiang Gao, Bingwen Zhang, Yubai Shi, Run-Wei Li, Zhicheng Zhong
Phys. Rev. Lett., 2025, 134 (7), 76101.
DOI: 10.1103/physrevlett.134.076101
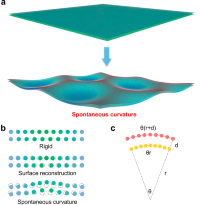
Spontaneous curvature in two-dimensional van der Waals heterostructures
Yuxiang Gao, Fenglin Deng, Ri He, Zhicheng Zhong
Nat. Commun., 2025, 16 (1), 717.
DOI: 10.1038/s41467-025-56055-x
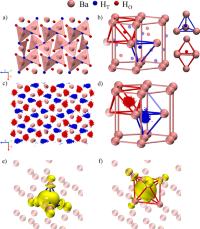
Machine learning-driven molecular dynamics unveils a bulk phase transformation driving ammonia synthesis on barium hydride
Axel Tosello Gardini, Umberto Raucci, Michele Parrinello
Nat. Commun., 2025, 16 (1), 2475.
DOI: 10.1038/s41467-025-57688-8
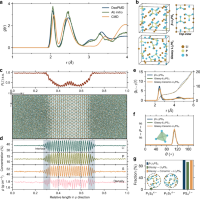
Disorder-induced enhancement of lithium-ion transport in solid-state electrolytes
Zhimin Chen, Tao Du, N. M. Anoop Krishnan, Yuanzheng Yue, Morten M. Smedskjaer
Nat. Commun., 2025, 16 (1), 1057.
DOI: 10.1038/s41467-025-56322-x
Harnessing Machine Learning to Enhance Transition State Search with Interatomic Potentials and Generative Models
Qiyuan Zhao, Yunhong Han, Duo Zhang, Jiaxu Wang, Peichen Zhong, Taoyong Cui, Bangchen Yin, Yirui Cao, Haojun Jia, Chenru Duan
Adv. Sci. (Weinheim Baden-Wurtt.- Ger.), 2025, 12 (34), e06240.
DOI: 10.1002/advs.202506240
Computational investigation of water glasses using machine-learning potentials
Ryan J. Szukalo, Nicolas Giovambattista, Pablo G. Debenedetti
Proc. National Acad. Sci. U. S. A., 2025, 122 (32), e2509609122.
DOI: 10.1073/pnas.2509609122

Short-range order stabilizes a cubic iron alloy in Earth's inner core
Zhi Li, Sandro Scandolo
Nat. Commun., 2025, 16 (1), 7574.
DOI: 10.1038/s41467-025-62666-1
Tuning water dissociation at oxide-electrolyte interfaces with electric fields
Chunyi Zhang, Zheng Yu, Roberto Car, Annabella Selloni
Proc. National Acad. Sci. U. S. A., 2025, 122 (34), e2505929122.
DOI: 10.1073/pnas.2505929122
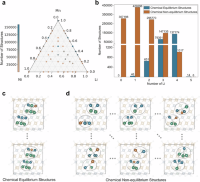
Dynamic oxygen-redox evolution of cathode reactions based on the multistate equilibrium potential model
Nian Ran, Chengbo Li, Qinwen Cui, Dezhen Xue, Jianjun Liu
npj Comput. Mater., 2025, 11 (1), 208.
DOI: 10.1038/s41524-025-01714-2
Unveiling the Lithium-Ion Transport Mechanism in Li 2 ZrCl 6 Solid-State Electrolyte via Deep Learning-Accelerated Molecular Dynamics Simulations
Hanzeng Guo, Volodymyr Koverga, Selva Chandrasekaran Selvaraj, Anh T. Ngo
Acs Appl. Energy Mater., 2025, 8 (19), 14773–14790.
DOI: 10.1021/acsaem.5c02491
Achieving Exceptionally Enhanced Thermal Conductivity and Bulk Modulus in Polar Insulators Via Modification of Chemical Bonding
Niraj Bhatt, Sandip Thakur, Pravin Karna, Ashutosh Giri
J. Phys. Chem. Lett., 2025, 16 (34), 8850–8860.
DOI: 10.1021/acs.jpclett.5c01476
Dynamic Metal-Support Interaction Dictates Cu Nanoparticle Sintering on Al2O3 Surfaces
Jiayan Xu, Shreeja Das, Amar Deep Pathak, Abhirup Patra, Sharan Shetty, Detlef Hohl, Roberto Car
ACS Nano, 2025, 19 (32), 29242–29254.
DOI: 10.1021/acsnano.5c04622
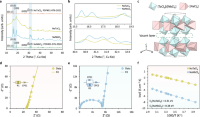
A sodium superionic chloride electrolyte driven by paddle wheel mechanism for solid state batteries
Rui Li, Kaiqi Xu, Shenhao Wen, Xiaohan Tang, Zheyu Lin, Xia Guo, Maxim Avdeev, Zhizhen Zhang, Yong-Sheng Hu
Nat. Commun., 2025, 16 (1), 6633.
DOI: 10.1038/s41467-025-61738-6

Interfaces govern the structure of angstrom-scale confined water solutions
Yongkang Wang, Fujie Tang, Xiaoqing Yu, Kuo-Yang Chiang, Chun-Chieh Yu, Tatsuhiko Ohto, Yunfei Chen, Yuki Nagata, Mischa Bonn
Nat. Commun., 2025, 16 (1), 7288.
DOI: 10.1038/s41467-025-62625-w
Developing a neural network machine learning interatomic potential for molecular dynamics simulations of La-Si-P systems
Ling Tang, Weiyi Xia, Gayatri Viswanathan, Ernesto Soto, Kirill Kovnir, Cai-Zhuang Wang
J. Chem. Phys., 2025, 163 (8), 84109.
DOI: 10.1063/5.0284672
Investigation of softer lattice dynamics in defect engineered GeTe crystals
Saptak Majumder, Pintu Singha, Sharath Kumar C, Mayanak K Gupta, Dharmendra Kumar, R Mittal, D K Shukla, M P Saravanan, Deepshikha Jaiswal-Nagar, Vinayak B Kamble
J. Phys. Condens. Matter, 2025, 37 (42), 425401.
DOI: 10.1088/1361-648x/ae0dd5

Predicting hydrogen diffusion in nickel-manganese random alloys using machine learning interatomic potentials
Kazuma Ito, Naoki Matsumura, Yuto Iwasaki, Yasufumi Sakai, Misaho Yamamura, Tomohiko Omura, Junichiro Yamabe, Hisao Matsunaga
Commun. Mater., 2025, 6 (1), 195.
DOI: 10.1038/s43246-025-00924-x

Learning atomic forces from uncertainty-calibrated adversarial attacks
Henrique Musseli Cezar, Tilmann Bodenstein, Henrik Andersen Sveinsson, Morten Ledum, Simen Reine, Sigbj\orn L\oland Bore
npj Comput. Mater., 2025, 11 (1), 200.
DOI: 10.1038/s41524-025-01703-5
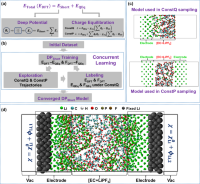
Observation of dendrite formation at Li metal--electrolyte interface by a machine-learning enhanced constant potential framework
Taiping Hu, Haichao Huang, Guobing Zhou, Xinyan Wang, Jiaxin Zhu, Zheng Cheng, Fangjia Fu, Xiaoxu Wang, Fuzhi Dai, Kuang Yu, Shenzhen Xu
Nature Communications, 2025, 16, 7379.
DOI: 10.1038/s41467-025-62824-5
Hexagonal ice density dependence on interatomic distance changes due to nuclear quantum effects
Lucas T. S. de Miranda, M'arcio S. Gomes-Filho, Mariana Rossi, Luana S. Pedroza, Alexandre R. Rocha
J. Chem. Phys., 2025, 163 (9), 94503.
DOI: 10.1063/5.0279956
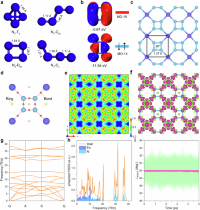
Machine learning augmented design of 2D magnet with planar cyclo- tetranitrogen: ambient thermal stability from quantum to mesoscale
Dong Fan, Ke Zheng, Hongfang Li, Junjie He, Pengbo Lyu
npj Comput. Mater., 2025, 11 (1), 286.
DOI: 10.1038/s41524-025-01763-7
Assessment and Application of Universal Machine Learning Interatomic Potentials in Solid-State Electrolyte Research
Hongwei Du, Xiang Huang, Jian Hui, Lanting Zhang, Yuanxun Zhou, Hong Wang
ACS Mater. Lett., 2025, 7 (10), 3403–3412.
DOI: 10.1021/acsmaterialslett.5c00336
DeePMD-kit v3: A multiple-backend framework for machine learning potentials
Jinzhe Zeng, Duo Zhang, Anyang Peng, Xiangyu Zhang, Sensen He, Yan Wang, Xinzijian Liu, Hangrui Bi, Yifan Li, Chun Cai, Chengqian Zhang, Yiming Du, Jia-Xin Zhu, Pinghui Mo, Zhengtao Huang, Qiyu Zeng, Shaochen Shi, Xuejian Qin, Zhaoxi Yu, Chenxing Luo, Ye Ding, Yun-Pei Liu, Ruosong Shi, Zhenyu Wang, Sigbjørn Løland Bore, Junhan Chang, Zhe Deng, Zhaohan Ding, Siyuan Han, Wanrun Jiang, Guolin Ke, Zhaoqing Liu, Denghui Lu, Koki Muraoka, Hananeh Oliaei, Anurag Kumar Singh, Haohui Que, Weihong Xu, Zhangmancang Xu, Yong-Bin Zhuang, Jiayu Dai, Timothy J. Giese, Weile Jia, Ben Xu, Darrin M. York, Linfeng Zhang, Han Wang
Journal of Chemical Theory and Computation, 2025, 21, 4375–4392.
DOI: 10.1021/acs.jctc.5c00340
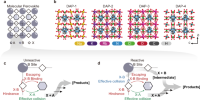
DeepEMs-25: a deep-learning potential to decipher kinetic tug-of-war dictating thermal stability in energetic materials
Ming-Yu Guo, Yun-Fan Yan, Pin Chen, Wei-Xiong Zhang
npj Computational Materials, 2025, 11 (1), 246.
DOI: 10.1038/s41524-025-01739-7
Deep potential-driven molecular dynamics of CO ice analogs: Investigating desorption following vibrational excitation
Maxime Infuso, Samuel Del Fré, Gilberto A. Alou Angulo, Mathieu Bertin, Jean-Hugues Fillion, Alejandro Rivero Santamaría, Maurice Monnerville
The Journal of Chemical Physics, 2025, 163 (8), 084303.
DOI: 10.1063/5.0285103
2024
Comparative analysis of ternary ( \textTi–Al–Nb ) interatomic potentials: moment tensor vs. deep learning approaches
Anju Chandran, Archa Santhosh, Claudio Pistidda, Paul Jerabek, Roland C. Aydin, Christian J. Cyron
Frontiers in Materials, 2024, 11.
DOI: 10.3389/fmats.2024.1466793
Software Infrastructure for Next-Generation QM/MM-$\Delta$MLP Force Fields
Timothy J. Giese, Jinzhe Zeng, Lauren Lerew, Erika McCarthy, Yujun Tao, \cS"olen Ekesan, Darrin M. York
J. Phys. Chem. B, 2024, 128 (26), 6257–6271.
DOI: 10.1021/acs.jpcb.4c01466
Amber free energy tools: Interoperable software for free energy simulations using generalized quantum mechanical/molecular mechanical and machine learning potentials
Yujun Tao, Timothy J. Giese, \cS"olen Ekesan, Jinzhe Zeng, B'alint Aradi, Ben Hourahine, Hasan Metin Aktulga, Andreas W. G"otz, Kenneth M. Merz Jr, Darrin M. York
J. Chem. Phys., 2024, 160 (22), 224104.
DOI: 10.1063/5.0211276
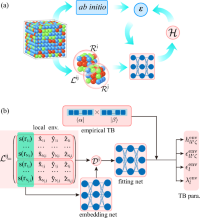
Deep learning tight-binding approach for large‐scale electronic simulations at finite temperatures with ab initio accuracy
Qiangqiang Gu, Zhanghao Zhouyin, Shishir Kumar Pandey, Peng Zhang, Linfeng Zhang, Weinan E
Nature Communications, 2024, 15 (1), 6772.
DOI: 10.1038/s41467-024-51006-4
Combining stochastic density functional theory with deep potential molecular dynamics to study warm dense matter
Tao Chen, Qianrui Liu, Yu Liu, Liang Sun, Mohan Chen
Matter Radiat. Extrem., 2024, 9 (1).
DOI: 10.1063/5.0163303
Double-Layer Distribution of Hydronium and Hydroxide Ions in the Air-Water Interface
Pengchao Zhang, Muye Feng, Xuefei Xu
ACS Phys. Chem Au, 2024.
DOI: 10.1021/acsphyschemau.3c00076
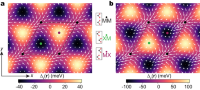
Polarization-driven band topology evolution in twisted MoTe2 and WSe2
Xiao-Wei Zhang, Chong Wang, Xiaoyu Liu, Yueyao Fan, Ting Cao, Di Xiao
Nat. Commun., 2024, 15 (1), 4223.
DOI: 10.1038/s41467-024-48511-x
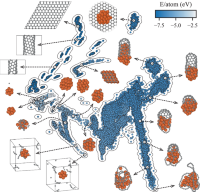
Dynamics of growing carbon nanotube interfaces probed by machine learning-enabled molecular simulations
Daniel Hedman, Ben McLean, Christophe Bichara, Shigeo Maruyama, J. Andreas Larsson, Feng Ding
Nat. Commun., 2024, 15 (1), 4076.
DOI: 10.1038/s41467-024-47999-7
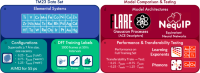
Complexity of many-body interactions in transition metals via machine- learned force fields from the TM23 data set
Cameron J. Owen, Steven B. Torrisi, Yu Xie, Simon Batzner, Kyle Bystrom, Jennifer Coulter, Albert Musaelian, Lixin Sun, Boris Kozinsky
npj Comput. Mater., 2024, 10 (1), 92.
DOI: 10.1038/s41524-024-01264-z
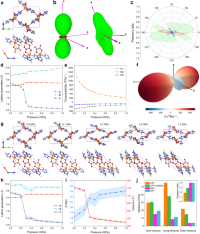
Unconventional mechanical and thermal behaviours of MOF CALF-20
Dong Fan, Supriyo Naskar, Guillaume Maurin
Nat. Commun., 2024, 15 (1), 3251.
DOI: 10.1038/s41467-024-47695-6
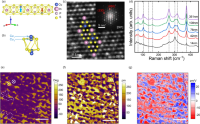
Realization of sextuple polarization states and interstate switching in antiferroelectric CuInP2S6
Tao Li, Yongyi Wu, Guoliang Yu, Shengxian Li, Yifeng Ren, Yadong Liu, Jiarui Liu, Hao Feng, Yu Deng, Mingxing Chen, Zhenyu Zhang, Tai Min
Nat. Commun., 2024, 15 (1), 2653.
DOI: 10.1038/s41467-024-46891-8
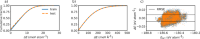
Unraveling the crystallization kinetics of the Ge2Sb2Te5 phase change compound with a machine-learned interatomic potential
Omar {Abou El Kheir}, Luigi Bonati, Michele Parrinello, Marco Bernasconi
npj Comput. Mater., 2024, 10 (1), 33.
DOI: 10.1038/s41524-024-01217-6
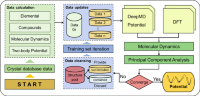
Principal component analysis enables the design of deep learning potential precisely capturing LLZO phase transitions
Yiwei You, Dexin Zhang, Fulun Wu, Xinrui Cao, Yang Sun, Zi-Zhong Zhu, Shunqing Wu
npj Comput. Mater., 2024, 10 (1), 57.
DOI: 10.1038/s41524-024-01240-7
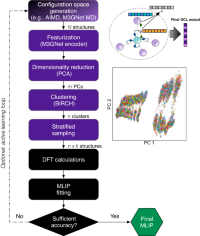
Robust training of machine learning interatomic potentials with dimensionality reduction and stratified sampling
Ji Qi, Tsz Wai Ko, Brandon C. Wood, Tuan Anh Pham, Shyue Ping Ong
npj Comput. Mater., 2024, 10 (1), 43.
DOI: 10.1038/s41524-024-01227-4
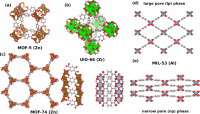
Machine learned force-fields for an Ab-initio quality description of metal-organic frameworks
Sandro Wieser, Egbert Zojer
npj Comput. Mater., 2024, 10 (1), 18.
DOI: 10.1038/s41524-024-01205-w
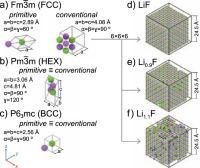
Enhancing ReaxFF for molecular dynamics simulations of lithium-ion batteries: an interactive reparameterization protocol
Paolo {De Angelis}, Roberta Cappabianca, Matteo Fasano, Pietro Asinari, Eliodoro Chiavazzo
Sci. Rep., 2024, 14 (1), 978.
DOI: 10.1038/s41598-023-50978-5
Ce-doped copper oxide and copper vanadate Cu3VO4 hybrid for boosting nitrate electroreduction to ammonia
Meng Zhang, Yang Liu, Yun Duan, Xu Liu, Yan-Qin Wang
J. Colloid Interface Sci., 2024, 671, 258–269.
DOI: 10.1016/j.jcis.2024.05.189
Construction and application of deep learning potential for CaO under high pressure
Xinwei Wang, Zi-Jiang Liu, Jin-Shan Feng, Meng-Ru Chen, Liang Li, Xiao-Wei Sun, Fubo Tian
Comput. Mater. Sci., 2024, 244, 113154.
DOI: 10.1016/j.commatsci.2024.113154
Pressure-driven enhancement of phonon contribution to the thermal conductivity of Iridium
Niraj Bhatt, Pravin Karna, Sandip Thakur, Ashutosh Giri
Int. J. Heat Mass Transf., 2024, 229, 125673.
DOI: 10.1016/j.ijheatmasstransfer.2024.125673
Selective activation of methane on hydroxyapatite surfaces: Insights from machine learning and density functional theory
Jing Wang, Xinrong Yan, Xin Wang, Mingli Yang, Dingguo Xu
Nano Energy, 2024, 127, 109762.
DOI: 10.1016/j.nanoen.2024.109762
Machine learning model to efficiently predict the structure and properties of MgCl2-NaCl-KCl melts
Taixi Feng, Jia Zhao, Guimin Lu
Sol. Energy Mater. Sol. Cells, 2024, 272, 112903.
DOI: 10.1016/j.solmat.2024.112903
Exploring dielectric properties in atomistic models of amorphous boron nitride
Thomas Galvani, Ali K Hamze, Laura Caputo, Onurcan Kaya, Simon M-M Dubois, Luigi Colombo, Viet-Hung Nguyen, Yongwoo Shin, Hyeon-Jin Shin, Jean-Christophe Charlier, Stephan Roche
J. Phys. Mater., 2024, 7 (3), 35003.
DOI: 10.1088/2515-7639/ad4c06
Dual-channel phonon transport leads to low thermal conductivity in pyrochlore La2Hf2O7
Junwei Che, Wenjie Huang, Guoliang Ren, Jiajun Linghu, Xuezhi Wang
Ceram. Int., 2024, 50 (13), 22865–22873.
DOI: 10.1016/j.ceramint.2024.04.011
Unveiling the crystallization mechanism of cadmium selenide via molecular dynamics simulation with machine-learning-based deep potential
Linshuang Zhang, Manyi Yang, Shiwei Zhang, Haiyang Niu
J. Mater. Sci. Technol., 2024, 185, 23–31.
DOI: 10.1016/j.jmst.2023.09.059
The nucleation and growth mechanism of solid-state amorphization and diffusion behavior at the W-Cu interface
Kai Wang, Guoqing Yao, Mengwei Lv, Zumin Wang, Yuan Huang, Wei Xi
Compos. B: Eng., 2024, 279, 111452.
DOI: 10.1016/j.compositesb.2024.111452
Machine learning molecular dynamics insight into high interface stability and fast kinetics of low-cost magnesium chloride amine electrolyte for rechargeable magnesium batteries
Haiming Hua, Fei Wang, Feng Wang, Jiayue Wu, Yaoqi Xu, Yichao Zhuang, Jing Zeng, Jinbao Zhao
Energy Storage Mater., 2024, 70, 103470.
DOI: 10.1016/j.ensm.2024.103470
Prediction of Cr(VI) and As(V) adsorption on goethite using hybrid surface complexation-machine learning model
Kai Chen, Chuling Guo, Chaoping Wang, Shoushi Zhao, Beiyi Xiong, Guining Lu, John R. Reinfelder, Zhi Dang
Water Res., 2024, 256, 121580.
DOI: 10.1016/j.watres.2024.121580
Thermal transports of 2D phosphorous carbides by machine learning molecular dynamics simulations
Chenyang Cao, Shuo Cao, YuanXu Zhu, Haikuan Dong, Yanzhou Wang, Ping Qian
Int. J. Heat Mass Transf., 2024, 224, 125359.
DOI: 10.1016/j.ijheatmasstransfer.2024.125359
Unraveling medium-range order and melting mechanism of ZIF-4 under high temperature
Zuhao Shi, Bin Liu, Yuanzheng Yue, Arramel Arramel, Neng Li
J. Am. Ceram. Soc., 2024, 107 (6), 3845–3856.
DOI: 10.1111/jace.19741
A deep-neural network potential to study transformation-induced plasticity in zirconia
Jin-Yu Zhang, Ga"el Huynh, Fu-Zhi Dai, Tristan Albaret, Shi-Hao Zhang, Shigenobu Ogata, David Rodney
J. Eur. Ceram. Soc., 2024, 44 (6), 4243–4254.
DOI: 10.1016/j.jeurceramsoc.2024.01.007
IR Spectroscopy of Carboxylate-Passivated Semiconducting Nanocrystals: Simulation and Experiment
Jakub K. Sowa, Danielle M. Cadena, Arshad Mehmood, Benjamin G. Levine, Sean T. Roberts, Peter J. Rossky
J. Phys. Chem. C, 2024, 128 (21), 8724–8731.
DOI: 10.1021/acs.jpcc.4c01988
Accelerating Computation of Acidity Constants and Redox Potentials for Aqueous Organic Redox Flow Batteries by Machine Learning Potential- Based Molecular Dynamics
Feng Wang, Zebing Ma, Jun Cheng
J. Am. Chem. Soc., 2024, 146 (21), 14566–14575.
DOI: 10.1021/jacs.4c01221
First-principles-based machine learning interatomic potential for molecular dynamics simulations of 2D lateral MoS2/WS2 heterostructures
Xiangjun Liu, Baolong Wang, Kun Jia, Quanjie Wang, Di Wang, Yucheng Xiong
J. Appl. Phys., 2024, 135 (20).
DOI: 10.1063/5.0201527
On-the-fly kinetic Monte Carlo simulations with neural network potentials for surface diffusion and reaction
Tomoko Yokaichiya, Tatsushi Ikeda, Koki Muraoka, Akira Nakayama
J. Chem. Phys., 2024, 160 (20), 204108.
DOI: 10.1063/5.0199240
Competing Reaction Mechanisms of Peptide Bond Formation in Water Revealed by Deep Potential Molecular Dynamics and Path Sampling
Rolf David, I\~naki Tu\~n'on, Damien Laage
J. Am. Chem. Soc., 2024, 146 (20), 14213–14224.
DOI: 10.1021/jacs.4c03445
Ultra-flat bands at large twist angles in group-V twisted bilayer materials
Zhi-Xiong Que, Shu-Zong Li, Bo Huang, Zhi-Xiong Yang, Wei- Bing Zhang
J. Chem. Phys., 2024, 160 (19), 194710.
DOI: 10.1063/5.0197757
Revealing the reconstruction mechanism of AgPd nanoalloys under fluorination based on a multiscale deep learning potential
Longfei Guo, Shuang Shan, Xiaoqing Liu, Wanxuan Zhang, Peng Xu, Fanzhe Ma, Zhen Li, Chongyang Wang, Junpeng Wang, Fuyi Chen
J. Chem. Phys., 2024, 160 (17), 174313.
DOI: 10.1063/5.0205616
Perspective: Atomistic simulations of water and aqueous systems with machine learning potentials
Amir Omranpour, Pablo {Montero De Hijes}, J"org Behler, Christoph Dellago
J. Chem. Phys., 2024, 160 (17), 170901.
DOI: 10.1063/5.0201241
Exploring the Relationship between Composition and Li-Ion Conductivity in the Amorphous Li-La-Zr-O System
Dexin Zhang, Yiwei You, Fulun Wu, Xinrui Cao, Tie-Yu L"u, Yang Sun, Zi-Zhong Zhu, Shunqing Wu
ACS Mater. Lett., 2024, 6 (5), 1849–1855.
DOI: 10.1021/acsmaterialslett.3c01558
Quasi-Classical Trajectory Calculation of Rate Constants Using an Ab Initio Trained Machine Learning Model (aML-MD) with Multifidelity Data
Zhiyu Shi, Aditya Dilip Lele, Ahren W. Jasper, Stephen J. Klippenstein, Yiguang Ju
J. Phys. Chem. A, 2024, 128 (17), 3449–3457.
DOI: 10.1021/acs.jpca.4c00750
Exploring Li-Ion Transport Properties of Li3TiCl6: A Machine Learning Molecular Dynamics Study
Selva Chandrasekaran Selvaraj, Volodymyr Koverga, Anh T. Ngo
J. Electrochem. Soc., 2024, 171 (5), 50544.
DOI: 10.1149/1945-7111/ad4ac9
Deep-learning interatomic potential for iron at extreme conditions
Zhi Li, Sandro Scandolo
Phys. Rev. B, 2024, 109 (18), 184108.
DOI: 10.1103/PhysRevB.109.184108
Revisiting the phase diagram and piezoelectricity of lead zirconate titanate from first principles
Yubai Shi, Ri He, Bingwen Zhang, Zhicheng Zhong
Phys. Rev. B, 2024, 109 (17), 174104.
DOI: 10.1103/PhysRevB.109.174104
Structural and mechanical properties of monolayer amorphous carbon and boron nitride
Xi Zhang, Yu-Tian Zhang, Yun-Peng Wang, Shiyu Li, Shixuan Du, Yu-Yang Zhang, Sokrates T. Pantelides
Phys. Rev. B, 2024, 109 (17), 174106.
DOI: 10.1103/PhysRevB.109.174106
Non-equilibrium nature of fracture determines the crack paths
Pengjie Shi, Shizhe Feng, Zhiping Xu
Extrem. Mech. Lett., 2024, 68, 102151.
DOI: 10.1016/j.eml.2024.102151
Understanding cellulose pyrolysis via ab initio deep learning potential field
Yuqin Xiao, Yuxin Yan, Hainam Do, Richard Rankin, Haitao Zhao, Ping Qian, Keke Song, Tao Wu, Cheng Heng Pang
Bioresour. Technol., 2024, 399, 130590.
DOI: 10.1016/j.biortech.2024.130590
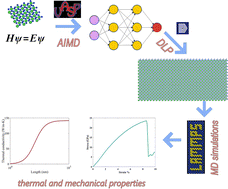
Lattice thermal conductivity and mechanical properties of the single- layer penta-NiN2 explored by a deep-learning interatomic potential
Pedram Mirchi, Christophe Adessi, Samy Merabia, Ali Rajabpour
Phys. Chem. Chem. Phys., 2024, 26 (19), 14216–14227.
DOI: 10.1039/d4cp00997e
Molecular dynamics simulations of heat transport using machine-learned potentials: A mini-review and tutorial on GPUMD with neuroevolution potentials
Haikuan Dong, Yongbo Shi, Penghua Ying, Ke Xu, Ting Liang, Yanzhou Wang, Zezhu Zeng, Xin Wu, Wenjiang Zhou, Shiyun Xiong, Shunda Chen, Zheyong Fan
J. Appl. Phys., 2024, 135 (16).
DOI: 10.1063/5.0200833
Neural network molecular dynamics study of LiGe2(PO4)3: Investigation of structure
I.A. Balyakin, M.I. Vlasov, S.V. Pershina, D.M. Tsymbarenko, A.A. Rempel
Comput. Mater. Sci., 2024, 239, 112979.
DOI: 10.1016/j.commatsci.2024.112979
Validation workflow for machine learning interatomic potentials for complex ceramics
Kimia Ghaffari, Salil Bavdekar, Douglas E. Spearot, Ghatu Subhash
Comput. Mater. Sci., 2024, 239, 112983.
DOI: 10.1016/j.commatsci.2024.112983
Deep Potential fitting and mechanical properties study of MgAlSi alloy
Chang-sheng Zhu, Wen-jing Dong, Zi-hao Gao, Li-jun Wang, Guang-zhao Li
Comput. Mater. Sci., 2024, 239, 112966.
DOI: 10.1016/j.commatsci.2024.112966
Investigation of the Degradation of LiPF6- in Polar Solvents through Deep Potential Molecular Dynamics
Da Zhu, Li Sheng, Taiping Hu, Sian Chen, Mengchao Shi, Haiming Hua, Kai Yang, Jianlong Wang, Yaping Tang, Xiangming He, Hong Xu
J. Phys. Chem. Lett., 2024, 15 (15), 4024–4030.
DOI: 10.1021/acs.jpclett.4c00575
Collaboration on Machine-Learned Potentials with IPSuite: A Modular Framework for Learning-on-the-Fly
Fabian Zills, Moritz Ren'e Sch"afer, Nico Segreto, Johannes K"astner, Christian Holm, Samuel Tovey
J. Phys. Chem. B, 2024, 128 (15), 3662–3676.
DOI: 10.1021/acs.jpcb.3c07187
Insights into Lithium Sulfide Glass Electrolyte Structures and Ionic Conductivity via Machine Learning Force Field Simulations
Rui Zhou, Kun Luo, Steve W. Martin, Qi An
ACS Appl. Mater. Interfaces, 2024, 16 (15), 18874–18887.
DOI: 10.1021/acsami.4c00618
Theoretical Study on Ion Diffusion Mechanism in W-Doped K3SbS4 as Solid-State Electrolyte for K-Ion Batteries
Rongyu Zhang, Shifeng Xu, Liyan Wang, Chuanyun Wang, Yongjun Zhou, Zhe L"u, Wenbo Li, Dan Xu, Sai Wang, Xu Yang
Inorg. Chem., 2024, 63 (15), 6743–6751.
DOI: 10.1021/acs.inorgchem.4c00074
Investigation of the effect of off-stoichiometric composition on oxygen transport in layered perovskite materials for SOFC cathode
Sung Hun Woo, Hyun Joo Yang, Yongseon Kim
Mater. Lett., 2024, 361, 136114.
DOI: 10.1016/j.matlet.2024.136114
Elucidating the local structure and properties of molten Na2CO3-K2CO3 salts using Machine Learning-Driven molecular dynamics
Taixi Feng, Bo Yang, Jia Zhao, Guimin Lu
Chem. Eng. Sci., 2024, 288, 119836.
DOI: 10.1016/j.ces.2024.119836
Development of machine learning force field for thermal conductivity analysis in MoAlB: Insights into anisotropic heat transfer mechanisms
Hanchao Zhang, Guoliang Ren, Peng Jia, Xiaofeng Zhao, Na Ni
Ceram. Int., 2024, 50 (8), 13740–13749.
DOI: 10.1016/j.ceramint.2024.01.288
Training machine learning potentials for reactive systems: A Colab tutorial on basic models
Xiaoliang Pan, Ryan Snyder, Jia-Ning Wang, Chance Lander, Carly Wickizer, Richard Van, Andrew Chesney, Yuanfei Xue, Yuezhi Mao, Ye Mei, Jingzhi Pu, Yihan Shao
J. Comput. Chem., 2024, 45 (10), 638–647.
DOI: 10.1002/jcc.27269
Unified deep learning network for enhanced accuracy in predicting thermal conductivity of bilayer graphene, hexagonal boron nitride, and their heterostructures
Rongkun Chen, Yu Tian, Jiayi Cao, Weina Ren, Shiqian Hu, Chunhua Zeng
J. Appl. Phys., 2024, 135 (14).
DOI: 10.1063/5.0201698
Many-body interactions and deep neural network potentials for water
Yaoguang Zhai, Richa Rashmi, Etienne Palos, Francesco Paesani
J. Chem. Phys., 2024, 160 (14), 144501.
DOI: 10.1063/5.0203682
Transferable Water Potentials Using Equivariant Neural Networks
Tristan Maxson, Tibor Szilv'asi
J. Phys. Chem. Lett., 2024, 15 (14), 3740–3747.
DOI: 10.1021/acs.jpclett.4c00605
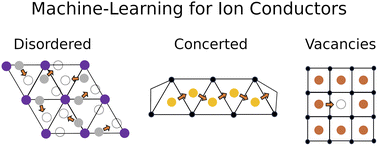
Accurate description of ion migration in solid-state ion conductors from machine-learning molecular dynamics
Takeru Miyagawa, Namita Krishnan, Manuel Grumet, Christian Rever'on Baecker, Waldemar Kaiser, David A. Egger
J. Mater. Chem. A, 2024, 12 (19), 11344–11361.
DOI: 10.1039/d4ta00452c
Effect of Interlayer Bonding on Superlubric Sliding of Graphene Contacts: A Machine-Learning Potential Study
Penghua Ying, Amir Natan, Oded Hod, Michael Urbakh
ACS Nano, 2024, 18 (14), 10133–10141.
DOI: 10.1021/acsnano.3c13099
Prediction of phonon properties of cubic boron nitride with vacancy defects and isotopic disorders by using a neural network potential
Jingwen Zhang, Junjie Zhang, Guoqiang Bao, Zehan Li, Xiaobo Li, Te-Huan Liu, Ronggui Yang
Appl. Phys. Lett., 2024, 124 (15).
DOI: 10.1063/5.0198431
Hydrocarbon Species on the Cu(111) Surface Studied with a Neural Network Potential
Sen Xu, Liling Wu, Yi Fan, Yufeng Liu, Xiongzhi Zeng, Zhenyu Li
J. Phys. Chem. C, 2024, 128 (13), 5697–5707.
DOI: 10.1021/acs.jpcc.3c08138
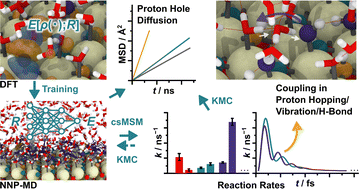
Long-range proton and hydroxide ion transfer dynamics at the water/CeO2 interface in the nanosecond regime: reactive molecular dynamics simulations and kinetic analysis
Taro Kobayashi, Tatsushi Ikeda, Akira Nakayama
Chem. Sci., 2024, 15 (18), 6816–6832.
DOI: 10.1039/d4sc01422g
Exploring diffusion behavior of superionic materials using machine- learning interatomic potentials
Cheng-Rong Hsing, Duc-Long Nguyen, Ching-Ming Wei
Phys. Rev. Mater., 2024, 8 (4), 43806.
DOI: 10.1103/PhysRevMaterials.8.043806
Photocatalytic activity of dual defect modified graphitic carbon nitride is robust to tautomerism: machine learning assisted ab initio quantum dynamics
Sraddha Agrawal, Bipeng Wang, Yifan Wu, David Casanova, Oleg V. Prezhdo
Nanoscale, 2024, 16 (18), 8986–8995.
DOI: 10.1039/d4nr00606b
Hydrogen Diffusion in the Lower Mantle Revealed by Machine Learning Potentials
Yihang Peng, Jie Deng
Jgr Solid Earth, 2024, 129 (4).
DOI: 10.1029/2023JB028333
Deciphering the controlling factors for phase transitions in zeolitic imidazolate frameworks
Tao Du, Shanwu Li, Sudheer Ganisetti, Mathieu Bauchy, Yuanzheng Yue, Morten M. Smedskjaer
Natl. Sci. Rev., 2024, 11 (4), nwae023.
DOI: 10.1093/nsr/nwae023
Machine learning heralding a new development phase in molecular dynamics simulations
Eva Pra\vsnikar, Martin Ljubi\vc, Andrej Perdih, Jure Bori\vsek
Artif Intell Rev, 2024, 57 (4), 102.
DOI: 10.1007/s10462-024-10731-4
Machine learning-based prediction of mechanical properties of N-doped $\gamma$-graphdiyne
Cun Zhang, Bolin Yang, Zhilong Peng, Shaohua Chen
Sci. China Mater., 2024, 67 (4), 1129–1139.
DOI: 10.1007/s40843-023-2733-7
Determining the thermal conductivity and phonon behavior of SiC materials with quantum accuracy via deep learning interatomic potential model
Baoqin Fu, Yandong Sun, Wanrun Jiang, Fu Wang, Linfeng Zhang, Han Wang, Ben Xu
J. Nucl. Mater., 2024, 591, 154897.
DOI: 10.1016/j.jnucmat.2024.154897
A 3D metallic porous sulfurized carbon anode identified by global structure search for Na-ion batteries with fast diffusion kinetics
Mohammed M. Obeid, Jiahui Liu, Penghu Du, Tongyu Liu, Qiang Sun
J. Energy Storage, 2024, 82, 110587.
DOI: 10.1016/j.est.2024.110587
Probing the critical point of MgSiO3 using deep potential simulation
Fei-Yang Xu, Zhi-Guo Li, Xiang-Rong Chen, Hua Y. Geng, Lei Liu, Jianbo Hu
J. Appl. Phys., 2024, 135 (12).
DOI: 10.1063/5.0189696
Ab Initio Driven Exploration on the Thermal Properties of Al-Li Alloy
Xiaoya Chang, Yongchao Wu, Qingzhao Chu, Gang Zhang, Dongping Chen
ACS Appl. Mater. Interfaces, 2024, 16 (12), 14954–14964.
DOI: 10.1021/acsami.4c01480
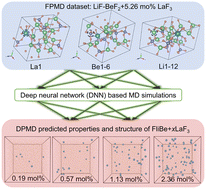
Compositional transferability of deep potential in molten LiF-BeF2 and LaF3 mixtures: prediction of density, viscosity, and local structure
Xuejiao Li, Tingrui Xu, Yu Gong
Phys. Chem. Chem. Phys., 2024, 26 (15), 12044–12052.
DOI: 10.1039/d4cp00079j
Compression Eliminates Charge Traps by Stabilizing Perovskite Grain Boundary Structures: An Ab Initio Analysis with Machine Learning Force Field
Dongyu Liu, Yifan Wu, Mikhail R. Samatov, Andrey S. Vasenko, Evgueni V. Chulkov, Oleg V. Prezhdo
Chem. Mater., 2024, 36 (6), 2898–2906.
DOI: 10.1021/acs.chemmater.3c03261
Neural Network-Based Sum-Frequency Generation Spectra of Pure and Acidified Water Interfaces with Air
Miguel {de la Puente}, Axel Gomez, Damien Laage
J. Phys. Chem. Lett., 2024, 15 (11), 3096–3102.
DOI: 10.1021/acs.jpclett.4c00113

A neural network potential energy surface assisted molecular dynamics study on the pyrolysis behavior of two spiro-hydrocarbons
Hang Xiao, Bin Yang
Phys. Chem. Chem. Phys., 2024, 26 (15), 11867–11879.
DOI: 10.1039/d3cp05425j
Thermal Conductivity of MgSiO3\ens uremath- H2O System Determined by Machine Learning Potentials
Yihang Peng, Jie Deng
Geophys. Res. Lett., 2024, 51 (5).
DOI: 10.1029/2023GL107245
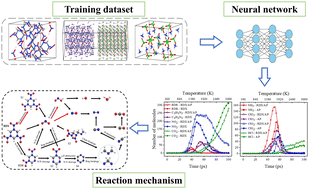
The thermal decomposition mechanism of RDX/AP composites: ab initio neural network MD simulations
Kehui Pang, Mingjie Wen, Xiaoya Chang, Yabei Xu, Qingzhao Chu, Dongping Chen
Phys. Chem. Chem. Phys., 2024, 26 (15), 11545–11557.
DOI: 10.1039/d3cp05709g
Electrical conductivity of copper under ultrahigh pressure and temperature conditions by both experiments and first-principles simulations
Bo Gan, Jun Li, Junjie Gao, Qiru Zeng, Wenhao Song, Yukai Zhuang, Yingxin Hua, Qiang Wu, Gang Jiang, Yuan Yin, Youjun Zhang
Phys. Rev. B, 2024, 109 (11), 115129.
DOI: 10.1103/PhysRevB.109.115129
A reactive molecular dynamics model for uranium/hydrogen containing systems
Artem Soshnikov, Rebecca Lindsey, Ambarish Kulkarni, Nir Goldman
J. Chem. Phys., 2024, 160 (9), 94117.
DOI: 10.1063/5.0183610
Deep Potential Molecular Dynamics Study of Propane Oxidative Dehydrogenation
Ziyi Liu, An-Hui Lu, Dongqi Wang
J. Phys. Chem. A, 2024, 128 (9), 1656–1664.
DOI: 10.1021/acs.jpca.3c07859
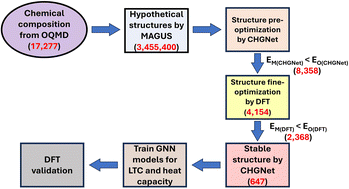
Graph theory and graph neural network assisted high-throughput crystal structure prediction and screening for energy conversion and storage
Joshua Ojih, Mohammed Al-Fahdi, Yagang Yao, Jianjun Hu, Ming Hu
J. Mater. Chem. A, 2024, 12 (14), 8502–8515.
DOI: 10.1039/d3ta06190f
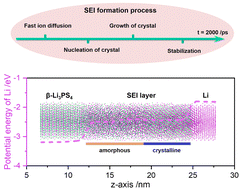
Visualizing the SEI formation between lithium metal and solid-state electrolyte
Fucheng Ren, Yuqi Wu, Wenhua Zuo, Wengao Zhao, Siyuan Pan, Hongxin Lin, Haichuan Yu, Jing Lin, Min Lin, Xiayin Yao, Torsten Brezesinski, Zhengliang Gong, Yong Yang
Energy Environ. Sci., 2024, 17 (8), 2743–2752.
DOI: 10.1039/d3ee03536k
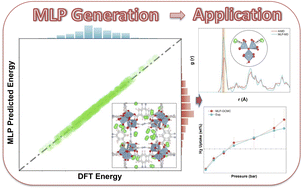
Machine learning potential for modelling H2 adsorption/diffusion in MOFs with open metal sites
Shanping Liu, Romain Dupuis, Dong Fan, Salma Benzaria, Mickaele Bonneau, Prashant Bhatt, Mohamed Eddaoudi, Guillaume Maurin
Chem. Sci., 2024, 15 (14), 5294–5302.
DOI: 10.1039/d3sc05612k
Modifying ring structures in lithium borate glasses under compression: MD simulations using a machine-learning potential
Shingo Urata, Aik Rui Tan, Rafael G'omez-Bombarelli
Phys. Rev. Mater., 2024, 8 (3), 33602.
DOI: 10.1103/PhysRevMaterials.8.033602
Thermoelastic properties of bridgmanite using deep-potential molecular dynamics
Tianqi Wan, Chenxing Luo, Yang Sun, Renata M. Wentzcovitch
Phys. Rev. B, 2024, 109 (9), 94101.
DOI: 10.1103/PhysRevB.109.094101
Understanding the dielectric relaxation of liquid water using neural network potential and classical pairwise potential
Jae Hyun Ryu, Ji Woong Yu, Tae Jun Yoon, Won Bo Lee
J. Mol. Liq., 2024, 397, 124054.
DOI: 10.1016/j.molliq.2024.124054
Synthetic pre-training for neural-network interatomic potentials
John L A Gardner, Kathryn T Baker, Volker L Deringer
Mach. Learn.: Sci. Technol., 2024, 5 (1), 15003.
DOI: 10.1088/2632-2153/ad1626
Review of deep learning algorithms in molecular simulations and perspective applications on petroleum engineering
Jie Liu, Tao Zhang, Shuyu Sun
Geosci. Front., 2024, 15 (2), 101735.
DOI: 10.1016/j.gsf.2023.101735
Machine-learned atomic cluster expansion potentials for fast and quantum-accurate thermal simulations of wurtzite AlN
Guang Yang, Yuan-Bin Liu, Lei Yang, Bing-Yang Cao
Arxiv, 2024, 135 (8).
DOI: 10.1063/5.0188905
Highly efficient and transferable interatomic potentials for $\alpha$-iron and $\alpha$-iron/hydrogen binary systems using deep neural networks
Shihao Zhang, Fanshun Meng, Rong Fu, Shigenobu Ogata
Comput. Mater. Sci., 2024, 235, 112843.
DOI: 10.1016/j.commatsci.2024.112843
Accurate and Transferable Machine Learning Potential for Molecular Dynamics Simulation of Sodium Silicate Glasses
Marco Bertani, Thibault Charpentier, Francesco Faglioni, Alfonso Pedone
J. Chem. Theory Comput., 2024, 20 (3), 1358–1370.
DOI: 10.1021/acs.jctc.3c01115
Development of a Neural Network Potential for Modeling Molten LiCl/KCl Salts: Bridging Efficiency and Accuracy
Abdullah {Bin Faheem}, Kyung-Koo Lee
J. Phys. Chem. C, 2024, 128 (5), 2163–2178.
DOI: 10.1021/acs.jpcc.3c07010
Screening Cu-Zeolites for Methane Activation Using Curriculum-Based Training
Jiawei Guo, Tyler Sours, Sam Holton, Chenghan Sun, Ambarish R. Kulkarni
ACS Catal., 2024, 14 (3), 1232–1242.
DOI: 10.1021/acscatal.3c05275
Strain-induced two-dimensional relaxor ferroelectrics in Se-doped PbTe
Zhanpeng Gong, Jefferson Zhe Liu, Xiangdong Ding, Jun Sun, Junkai Deng
Phys. Rev. B, 2024, 109 (5), 54117.
DOI: 10.1103/PhysRevB.109.054117
Effects of four-phonon interaction and vacancy defects on the thermal conductivity of the low-temperature phase of SnSe
Pan Zhang, Dan Jin, Mi Qin, Zhenhua Zhang, Yong Liu, Ziyu Wang, Zhihong Lu, Rui Xiong, Jing Shi
Phys. Rev. Appl., 2024, 21 (2), 24043.
DOI: 10.1103/PhysRevApplied.21.024043
Implementation and Validation of an OpenMM Plugin for the Deep Potential Representation of Potential Energy
Ye Ding, Jing Huang
Int. J. Mol. Sci., 2024, 25 (3), 1448.
DOI: 10.3390/ijms25031448
Synthesis and magnetic properties of some monotectic composites containing ultra-dispersed particles of YGdTbDyHo high-entropy alloy
S.A. Uporov, E.V. Sterkhov, I.A. Balyakin, V.A. Bykov, I.S. Sipatov, A.A. Rempel
Intermetallics, 2024, 165, 108121.
DOI: 10.1016/j.intermet.2023.108121
Combining Force Fields and Neural Networks for an Accurate Representation of Bonded Interactions
Ganesh Kamath, Alexey Illarionov, Serzhan Sakipov, Leonid Pereyaslavets, Igor V. Kurnikov, Oleg Butin, Ekaterina Voronina, Ilya Ivahnenko, Igor Leontyev, Grzegorz Nawrocki, Mikhail Darkhovskiy, Michael Olevanov, Yevhen K. Cherniavskyi, Christopher Lock, Sean Greenslade, YuChun Chen, Roger D. Kornberg, Michael Levitt, Boris Fain
J. Phys. Chem. A, 2024, 128 (4), 807–812.
DOI: 10.1021/acs.jpca.3c07598
Grain refinement mechanism of boron addition within Ti-Al alloy
B. Zhai, J. Chang, G. X. Li, H. P. Wang
Appl. Phys. A, 2024, 130 (2), 106.
DOI: 10.1007/s00339-024-07277-1
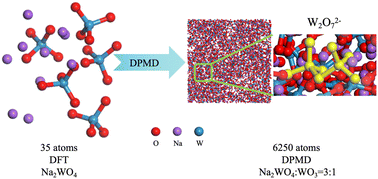
Effect of electric fields on tungsten distribution in Na2WO4-WO3 molten salt
Yuliang Guo, Xiaobo Sun, Handong Jiao, Liwen Zhang, Wenxuan Qin, Xiaoli Xi, Zuoren Nie
Phys. Chem. Chem. Phys., 2024, 26 (8), 6590–6599.
DOI: 10.1039/d3cp06202c
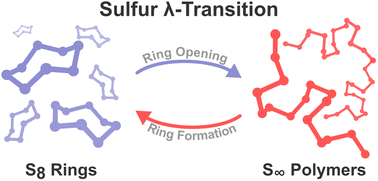
Structure and polymerization of liquid sulfur across the $\lambda$-transition
Manyi Yang, Enrico Trizio, Michele Parrinello
Chem. Sci., 2024, 15 (9), 3382–3392.
DOI: 10.1039/d3sc06282a
Deep learning interatomic potential for thermal and defect behaviour of aluminum nitride with quantum accuracy
Tao Li, Qing Hou, Jie-chao Cui, Jia-hui Yang, Ben Xu, Min Li, Jun Wang, Bao-qin Fu
Comput. Mater. Sci., 2024, 232, 112656.
DOI: 10.1016/j.commatsci.2023.112656
Illuminating the mechanical responses of amorphous boron nitride through deep learning: A molecular dynamics study
Shin-Pon Ju, Chao-Chuan Huang, Hsing-Yin Chen
Comput. Mater. Sci., 2024, 232, 112664.
DOI: 10.1016/j.commatsci.2023.112664
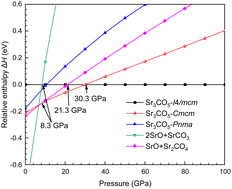
Study of high-pressure thermophysical properties of orthocarbonate Sr3CO5 using deep learning molecular dynamics simulations
Xin-Xuan Wang, Ting Song, Zhen-Shuai Lei, Xiao-Wei Sun, Jun-Hong Tian, Zi-Jiang Liu
Phys. Chem. Chem. Phys., 2024, 26 (7), 6351–6361.
DOI: 10.1039/d3cp04833k
Vacancy-induced phonon localization in boron arsenide using a unified neural network interatomic potential
Junjie Zhang, Hao Zhang, Jing Wu, Xin Qian, Bai Song, Cheng-Te Lin, Te-Huan Liu, Ronggui Yang
Cell Rep. Phys. Sci., 2024, 5 (1), 101760.
DOI: 10.1016/j.xcrp.2023.101760
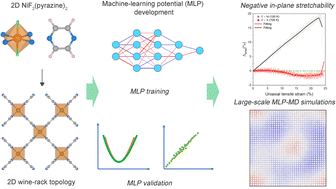
Unravelling abnormal in-plane stretchability of two-dimensional metal- organic frameworks by machine learning potential molecular dynamics
Dong Fan, Aydin Ozcan, Pengbo Lyu, Guillaume Maurin
Nanoscale, 2024, 16 (7), 3438–3447.
DOI: 10.1039/d3nr05966a
Hydration MgCl2-NaCl-KCl molten salt using a novel approach for training machine learning potential
Taixi Feng, Guimin Lu
J. Mol. Liq., 2024, 394, 123533.
DOI: 10.1016/j.molliq.2023.123533
Nuclear quantum effects in the acetylene:ammonia plastic co-crystal
Atul C. Thakur, Richard C. Remsing
J. Chem. Phys., 2024, 160 (2), 24502.
DOI: 10.1063/5.0179161
Molecular Simulation of Lithium Carbonate Reactive Vapor-Liquid Equilibria Using a Deep Potential Model
Dina Kussainova, Athanassios Z. Panagiotopoulos
J. Chem. Eng. Data, 2024, 69 (1), 204–214.
DOI: 10.1021/acs.jced.3c00580
Development of NaCl-MgCl2-CaCl2 Ternary Salt for High-Temperature Thermal Energy Storage Using Machine Learning
Wenhao Dong, Heqing Tian, Wenguang Zhang, Jun-Jie Zhou, Xinchang Pang
ACS Appl. Mater. Interfaces, 2024, 16 (1), 530–539.
DOI: 10.1021/acsami.3c13412
TrIP-Transformer Interatomic Potential Predicts Realistic Energy Surface Using Physical Bias
Bryce E. Hedelius, Damon Tingey, Dennis {Della Corte}
J. Chem. Theory Comput., 2024, 20 (1), 199–211.
DOI: 10.1021/acs.jctc.3c00936
Confinement Effects on Proton Transfer in TiO2 Nanopores from Machine Learning Potential Molecular Dynamics Simulations
Hyuna Kwon, Marcos F. {Calegari Andrade}, Shane Ardo, Daniel V. Esposito, Tuan Anh Pham, Tadashi Ogitsu
ACS Appl. Mater. Interfaces, 2024, 16 (24), 31687–31695.
DOI: 10.1021/acsami.4c02339
An ab- initio deep neural network potential for accurate large-scale simulations of the muscovite mica-water interface
Abhinav S. Raman, Annabella Selloni
Mol. Phys., 2024.
DOI: 10.1080/00268976.2024.2365430
Applications of machine-learning interatomic potentials for modeling ceramics, glass, and electrolytes: A review
Shingo Urata, Marco Bertani, Alfonso Pedone
J. Am. Ceram. Soc., 2024.
DOI: 10.1111/jace.19934
Accelerating Molecular Vibrational Spectra Simulations with a Physically Informed Deep Learning Model
Yuzhuo Chen, Sebastian V. Pios, Maxim F. Gelin, Lipeng Chen
J. Chem. Theory Comput., 2024, 20 (11), 4703–4710.
DOI: 10.1021/acs.jctc.4c00173
Structural Origin of the Deformation Propensity of Zeolitic Imidazolate Framework Glasses
Tao Du, Xuan Ge, Fengming Cao, Han Liu, Caijuan Shi, Junwei Ding, Daming Sun, Qiangqiang Zhang, Yuanzheng Yue, Morten M. Smedskjaer
Chem. Mater., 2024, 36 (12), 6167–6179.
DOI: 10.1021/acs.chemmater.4c00921
Phase Transition in Silicon from Machine Learning Informed Metadynamics
Mangladeep Bhullar, Zihao Bai, Akinwumi Akinpelu, Yansun Yao
ChemPhysChem, 2024, e202400090.
DOI: 10.1002/cphc.202400090
The structural evolution of SiBCNZr amorphous ceramics analyzed by machine learning potential
Meng Zhang, Siyan Deng, Jianghong Zhang, Daxin Li, Dechang Jia, Yu Zhou
Ceram. Int., 2024, 50 (17), 30008–30017.
DOI: 10.1016/j.ceramint.2024.05.296
Machine Learning Molecular Dynamics Shows Anomalous Entropic Effect on Catalysis through Surface Pre-melting of Nanoclusters
Fu-Qiang Gong, Yun-Pei Liu, Ye Wang, Weinan E, Zhong-Qun Tian, Jun Cheng
Angew. Chem. (International, Engl.), 2024, 63 (27), e202405379.
DOI: 10.1002/anie.202405379
Deep potential molecular dynamic and electrochemical experiments to reveal the structure and behavior of Mn(II) in magnesium electrolysis
Taixi Feng, Zhaoting Liu, Guimin Lu
Braz. J. Chem. Eng., 2024.
DOI: 10.1007/s43153-024-00465-9
Stacked High-Entropy Hydroxides Promote Charge Transfer Kinetics for Photoelectrochemical Water Splitting
Lei Wang, Zehua Gao, Kerong Su, Nhat Truong Nguyen, Rui-Ting Gao, Junxiang Chen, Lei Wang
Adv. Funct. Mater., 2024.
DOI: 10.1002/adfm.202403948
Improving the reliability of machine learned potentials for modeling inhomogeneous liquids
Kamron Fazel, Nima Karimitari, Tanooj Shah, Christopher Sutton, Ravishankar Sundararaman
J. Comput. Chem., 2024, 45 (21), 1821–1828.
DOI: 10.1002/jcc.27353
Fast Room-Temperature Mg-Ion Conduction in Clay-Like Halide Glassy Electrolytes
Xiaochen Yang, Sunny Gupta, Yu Chen, Dogancan Sari, Han-Ming Hau, Zijian Cai, Chaochao Dun, Miao Qi, Lu Ma, Yi Liu, Jeffrey J. Urban, Gerbrand Ceder
Adv. Energy Mater., 2024.
DOI: 10.1002/aenm.202400163
Amorphous solidification mechanism of multicomponent Ti-Cu based alloy investigated by molecular dynamics simulation and drop tube experiments
ShanSen XU, Jian CHANG, Bin ZHAI, PengXu YAN, MaoJie LIN, BingBo WEI
Sci. Sin.-Tech., 2024, 54 (3), 477–489.
DOI: 10.1360/SST-2023-0408
Impact of Amorphous LiF Coating Layers on Cathode-Electrolyte Interfaces in Solid-State Batteries
Taiping Hu, Linhan Xu, Fuzhi Dai, Guobing Zhou, Fangjia Fu, Xiaoxu Wang, Linsen Li, Xinping Ai, Shenzhen Xu
Adv. Funct. Mater., 2024.
DOI: 10.1002/adfm.202402993
Prediction of Mechanical Properties of High-Entropy Carbide (Ti0.2Zr0.2Hf0.2Nb0.2Ta0.2)C with the Use of Machine Learning Potential
N. S. Pikalova, I. A. Balyakin, A. A. Yuryev, A. A. Rempel
Dokl Phys Chem, 2024, 514 (1), 9–14.
DOI: 10.1134/S0012501624600049
Hyper-Elastic Deformation via Martensitic Phase Transformation in Cadmium Telluride
Kun Luo, Xiao Han, Jonathan Cappola, Dian Li, Yufeng Zheng, Lin Li, Feng Yan, Qi An
Adv Eng Mater, 2024.
DOI: 10.1002/adem.202302076
Probing the state of hydrogen in $\delta$-AlOOH at mantle conditions with machine learning potential
Chenxing Luo, Yang Sun, Renata M. Wentzcovitch
Phys. Rev. Res., 2024, 6 (1), 13292.
DOI: 10.1103/PhysRevResearch.6.013292
Ultralow-Friction at Cryogenic Temperature Induced by Hydrogen Correlated Quantum Effect
Weiqi Chen, Kang Wang, Xinran Miao, Jie Zhang, Aisheng Song, Xinchun Chen, Jianbin Luo, Tianbao Ma
Small (Weinheim Bergstr. Ger.), 2024, e2400083.
DOI: 10.1002/smll.202400083
Thermal transport of glasses via machine learning driven simulations
Paolo Pegolo, Federico Grasselli
Front. Mater., 2024, 11.
DOI: 10.3389/fmats.2024.1369034
Determining the mechanical and decomposition properties of high energetic materials ($\alpha$-RDX, $\beta$-HMX, and $\varepsilon$-CL-20) using a neural network potential
Mingjie Wen, Xiaoya Chang, Yabei Xu, Dongping Chen, Qingzhao Chu
Phys. Chem. Chem. Phys., 2024, 26 (13), 9984–9997.
DOI: 10.1039/d4cp00017j
A Perspective on the Molecular Modeling of Electrolyte Decomposition Reactions for Solid Electrolyte Interphase Growth in Lithium-Ion Batteries
Mohammed {Bin Jassar}, Carine Michel, Sara Abada, Theodorus {De Bruin}, Sylvain Tant, Carlos Nieto-Draghi, Stephan N. Steinmann
Adv. Funct. Mater., 2024.
DOI: 10.1002/adfm.202313188
Tunable Ferroelectric Topological Defects on 2D Topological Surfaces: Complex Strain Engineering Skyrmion-Like Polar Structures in 2D Materials
Bo Xu, Zhanpeng Gong, Jingran Liu, Yunfei Hong, Yang Yang, Lou Li, Yilun Liu, Junkai Deng, Jefferson Zhe Liu
Adv. Funct. Mater., 2024, 34 (26).
DOI: 10.1002/adfm.202311599
Theoretical Prediction of Thermal Expansion Anisotropy for Y2Si2O7 Environmental Barrier Coatings Using a Deep Neural Network Potential and Comparison to Experiment
Cameron J. Bodenschatz, Wissam A. Saidi, Jamesa L. Stokes, Rebekah I. Webster, Gustavo Costa
Mater. (Basel Switz.), 2024, 17 (2), 286.
DOI: 10.3390/ma17020286
Accumulative coarse-graining of simple structural descriptors enables accurately predicting the dynamics of metallic liquids
Jiang You, Cheng Wang, Qi Wang, Min Zha, Hai-Long Jia, Jian Wang, Hui-Yuan Wang
Mater. Today Phys., 2024, 40, 101282.
DOI: 10.1016/j.mtphys.2023.101282
Deep machine learning, molecular dynamics and experimental studies of liquid Al-Cu-Co alloys
L.V. Kamaeva, E.N. Tsiok, N.M. Chtchelkatchev
J. Mol. Liq., 2024, 393, 123659.
DOI: 10.1016/j.molliq.2023.123659
Development of U-Zr-Xe ternary interatomic potentials appropriate for simulation of defect and Xe behaviors in U-Zr system
Peng Jiang, Rongyang Qiu, Jinli Cao, Xichuan Liao, Yangchun Chen, Zhixiao Liu, Xinfu He, Wen Yang, Huiqiu Deng
J. Nucl. Mater., 2024, 588, 154824.
DOI: 10.1016/j.jnucmat.2023.154824
Active learning molecular dynamics-assisted insights into ultralow thermal conductivity of two-dimensional covalent organic frameworks
Zhiqiang Li, Haoyu Dong, Jian Wang, Linhua Liu, Jia-Yue Yang
Int. J. Heat Mass Transf., 2024, 225, 125404.
DOI: 10.1016/j.ijheatmasstransfer.2024.125404
Prebiotic chemical reactivity in solution with quantum accuracy and microsecond sampling using neural network potentials
Zakarya Benayad, Rolf David, Guillaume Stirnemann
Proc. Natl. Acad. Sci. U. S. A., 2024, 121 (23), e2322040121.
DOI: 10.1073/pnas.2322040121
Neural network potential-based molecular investigation of pollutant formation of ammonia and ammonia-hydrogen combustion
Zhihao Xing, Xi Jiang
Chem. Eng. J., 2024, 489, 151492.
DOI: 10.1016/j.cej.2024.151492
Weaving the next-level structure of calcium silicate hydrate at the submicron scale via a remapping algorithm from coarse-grained to all- atom model
Ao Zhou, Juntao Kang, Renyuan Qin, Huali Hao, Tiejun Liu, Zechuan Yu
Cem. Concr. Res., 2024, 180, 107501.
DOI: 10.1016/j.cemconres.2024.107501
Ab initio dispersion potentials based on physics-based functional forms with machine learning
Corentin Villot, Ka Un Lao
J. Chem. Phys., 2024, 160 (18), 184103.
DOI: 10.1063/5.0204064
Homogeneous water vapor condensation with a deep neural network potential model
Shenghui Zhong, Zheyu Shi, Bin Zhang, Zhengcheng Wen, Longfei Chen
J. Chem. Phys., 2024, 160 (12), 124303.
DOI: 10.1063/5.0189448
Machine learning is funny but physics makes the money: How machine- learning potentials can advance computer-aided materials design in metallurgy
M. Hodapp
Comput. Mater. Sci., 2024, 233, 112715.
DOI: 10.1016/j.commatsci.2023.112715
DP/MM: A Hybrid Model for Zinc-Protein Interactions in Molecular Dynamics
Ye Ding, Jing Huang
J. Phys. Chem. Lett., 2024, 15 (2), 616–627.
DOI: 10.1021/acs.jpclett.3c03158
Calculation of solvation force in molecular dynamics simulation by deep-learning method
Jun Liao, Mincong Wu, Junyong Gao, Changjun Chen
Biophys. J., 2024, S0006-3495(24)00163-2.
DOI: 10.1016/j.bpj.2024.02.029
Ultrafast switching dynamics of the ferroelectric order in stacking- engineered ferroelectrics
Ri He, Bingwen Zhang, Hua Wang, Lei Li, Ping Tang, Gerrit Bauer, Zhicheng Zhong
Acta Materialia, 2024, 262, 119416.
DOI: 10.1016/j.actamat.2023.119416
Unraveling pyrolysis mechanisms of lignin dimer model compounds: Neural network-based molecular dynamics simulation investigations
Zhe Shang, Hui Li
Fuel, 2024, 357, 129909.
DOI: 10.1016/j.fuel.2023.129909
Mobile dislocation mediated Hall-Petch and inverse Hall-Petch behaviors in nanocrystalline Al-doped boron carbide
Jun Li, Kun Luo, Qi An
Journal of the European Ceramic Society, 2024, 44, 659–667.
DOI: 10.1016/j.jeurceramsoc.2023.09.079
2023
Data Efficient and Stability Indicated Sampling for Developing Reactive Machine Learning Potential to Achieve Ultralong Simulation in Lithium-Metal Batteries
Longkun Xu, Wei Shao, Haishun Jin, Qiang Wang
J. Phys. Chem. C, 2023, 127 (50), 24106–24117.
DOI: 10.1021/acs.jpcc.3c05522
Ultralow Lattice Thermal Transport and Considerable Wave-like Phonon Tunneling in Chalcogenide Perovskite BaZrS3
Yu Wu, Ying Chen, Zhenxing Fang, Yimin Ding, Qiaoqiao Li, Kui Xue, Hezhu Shao, Hao Zhang, Liujiang Zhou
J. Phys. Chem. Lett., 2023, 14 (50), 11465–11473.
DOI: 10.1021/acs.jpclett.3c02940
Microstructure determination of PdRu immiscible alloys based on electron-pair distribution function and local elemental segregation
Zheng Hu, Hui Li, Wenbo Zhao, Wei Zhou, Shi Hu
Cell Rep. Phys. Sci., 2023, 4 (12), 101713.
DOI: 10.1016/j.xcrp.2023.101713
Coarse-Graining with Equivariant Neural Networks: A Path Toward Accurate and Data-Efficient Models
Timothy D. Loose, Patrick G. Sahrmann, Thomas S. Qu, Gregory A. Voth
J. Phys. Chem. B, 2023, 127 (49), 10564–10572.
DOI: 10.1021/acs.jpcb.3c05928
Ferroelectric Domain and Switching Dynamics in Curved In2Se3: First- Principles and Deep Learning Molecular Dynamics Simulations
Dongyu Bai, Yihan Nie, Jing Shang, Junxian Liu, Minghao Liu, Yang Yang, Haifei Zhan, Liangzhi Kou, Yuantong Gu
Nano Lett., 2023, 23 (23), 10922–10929.
DOI: 10.1021/acs.nanolett.3c03160
Efficient Molecular Dynamics Simulations of Deep Eutectic Solvents with First-Principles Accuracy Using Machine Learning Interatomic Potentials
Omid Shayestehpour, Stefan Zahn
J. Chem. Theory Comput., 2023, 19 (23), 8732–8742.
DOI: 10.1021/acs.jctc.3c00944
Pressure-induced one-dimensional oxygen ion diffusion channel in lead apatite
Ri He, Hongyu Wu, Xuejian Qin, Xuejiao Chen, Zhicheng Zhong
Phys. Rev. B, 2023, 108 (22), 224114.
DOI: 10.1103/PhysRevB.108.224114
Machine learning of microscopic structure-dynamics relationships in complex molecular systems
Martina Crippa, Annalisa Cardellini, Matteo Cioni, G'abor Cs'anyi, Giovanni M Pavan
Mach. Learn.: Sci. Technol., 2023, 4 (4), 45044.
DOI: 10.1088/2632-2153/ad0fa5
Unlocking the mysterious polytypic features within vaterite CaCO3
Xingyuan San, Junwei Hu, Mingyi Chen, Haiyang Niu, Paul J. M. Smeets, Christos D. Malliakas, Jie Deng, Kunmo Koo, Roberto {Dos Reis}, Vinayak P. Dravid, Xiaobing Hu
Nat. Commun., 2023, 14 (1), 7858.
DOI: 10.1038/s41467-023-43625-0
Full-scale ab initio simulations of laser-driven atomistic dynamics
Qiyu Zeng, Bo Chen, Shen Zhang, Dongdong Kang, Han Wang, Xiaoxiang Yu, Jiayu Dai
npj Comput. Mater., 2023, 9 (1), 213.
DOI: 10.1038/s41524-023-01168-4
Double superionicity in icy compounds at planetary interior conditions
Kyla {de Villa}, Felipe Gonz'alez-Cataldo, Burkhard Militzer
Nat. Commun., 2023, 14 (1), 7580.
DOI: 10.1038/s41467-023-42958-0
Microstructure evolution under thermo-mechanical operating of rocksalt-structure TiN via neural network potential
Fangyu Guo, Bo Chen, Qiyu Zeng, Xiaoxiang Yu, Kaiguo Chen, Dongdong Kang, Yong Du, Jianhua Wu, Jiayu Dai
J. Chem. Phys., 2023, 159 (20), 204702.
DOI: 10.1063/5.0171528
Molecular Dynamics Study of Silicon Carbide Using an Ab Initio-Based Neural Network Potential: Effect of Composition and Temperature on Crystallization Behavior
Jinyoung Lim, Youngseon Shim, Jaehong Park, Hongkee Yoon, Munbo Shim, Young-Gu Kim, Dae Sin Kim
J. Phys. Chem. C, 2023, 127 (46), 22692–22703.
DOI: 10.1021/acs.jpcc.3c04224
How the Acidity of Water Droplets and Films Is Controlled by the Air- Water Interface
Miguel {de la Puente}, Damien Laage
J. Am. Chem. Soc., 2023, 145 (46), 25186–25194.
DOI: 10.1021/jacs.3c07506
Tutorial: Deep learning prediction of thermophysical properties for liquid multicomponent alloys
R. L. Xiao, K. L. Liu, Y. Ruan, L. Hu, B. Wei
J. Appl. Phys., 2023, 134 (19).
DOI: 10.1063/5.0173250
Exploring the formation of gold/silver nanoalloys with gas-phase synthesis and machine-learning assisted simulations
Quentin Gromoff, Patrizio Benzo, Wissam A. Saidi, Christopher M. Andolina, Marie-Jos'e Casanove, Teresa Hungria, Sophie Barre, Magali Benoit, Julien Lam
Nanoscale, 2023, 16 (1), 384–393.
DOI: 10.1039/d3nr04471h
Modeling Exchange Reactions in Covalent Adaptable Networks with Machine Learning Force Fields
Yaguang Sun, Kaiwei Wan, Wenhui Shen, Jianxin He, Tong Zhou, Hui Wang, Hua Yang, Xinghua Shi
Macromolecules, 2023, 56 (21), 9003–9013.
DOI: 10.1021/acs.macromol.3c01377
Transferable Force Field for Gallium Nitride Crystal Growth from the Melt Using On-The-Fly Active Learning
Xiangyu Chen, William Shao, Nam Q. Le, Paulette Clancy
J. Chem. Theory Comput., 2023, 19 (21), 7861–7872.
DOI: 10.1021/acs.jctc.3c00587
Anomalous Thermal Transport across the Superionic Transition in Ice
Rong Rong {Qiu Qiu }, Qiyu Qi Yu {Zeng Ceng }, Han Han {Wang Wang }, Dongdong Dong Dong {Kang Kang }, Xiaoxiang Xiao Xiang {Yu Yu }, Jiayu Jia Yu {Dai Dai }
Chin. Phys. Lett., 2023, 40 (11), 116301.
DOI: 10.1088/0256-307X/40/11/116301
Universal interatomic potential for perovskite oxides
Jing Wu, Jiyuan Yang, Yuan-Jinsheng Liu, Duo Zhang, Yudi Yang, Yuzhi Zhang, Linfeng Zhang, Shi Liu
Phys. Rev. B, 2023, 108 (18), L180104.
DOI: 10.1103/PhysRevB.108.L180104
Influence of temperature on the process of hydrogen bond symmetrization in $\varepsilon$- FeOOH
Zhen-Shuai Lei, Xiao-Wei Sun, Xin-Xuan Wang, Zi-Jiang Liu, Ting Song, Jun-Hong Tian
Phys. Rev. B, 2023, 108 (18), 184105.
DOI: 10.1103/PhysRevB.108.184105
Transition from electron-dominated to phonon-driven thermal transport in tungsten under extreme pressures
Niraj Bhatt, Pravin Karna, Sandip Thakur, Ashutosh Giri
Phys. Rev. Mater., 2023, 7 (11), 115001.
DOI: 10.1103/PhysRevMaterials.7.115001
Strain-aided room-temperature second-order ferroelectric phase transition in monolayer PbTe: Deep potential molecular dynamics simulations
Zhanpeng Gong, Jefferson Zhe Liu, Xiangdong Ding, Jun Sun, Junkai Deng
Phys. Rev. B, 2023, 108 (13), 134112.
DOI: 10.1103/PhysRevB.108.134112
The role of dynamics in heterogeneous catalysis: Surface diffusivity and N2 decomposition on Fe(111)
Luigi Bonati, Daniela Polino, Cristina Pizzolitto, Pierdomenico Biasi, Rene Eckert, Stephan Reitmeier, Robert Schl"ogl, Michele Parrinello
Proc. Natl. Acad. Sci. U. S. A., 2023, 120 (50), e2313023120.
DOI: 10.1073/pnas.2313023120
Active learning prediction and experimental confirmation of atomic structure and thermophysical properties for liquid Hf_{76}W_{24} refractory alloy
K. L. Liu, R. L. Xiao, Y. Ruan, B. Wei
Phys. Rev. E, 2023, 108 (5-2), 55310.
DOI: 10.1103/PhysRevE.108.055310
Machine learning interatomic potential for molecular dynamics simulation of the ferroelectric KNbO3 perovskite
Hao-Cheng Thong, XiaoYang Wang, Jian Han, Linfeng Zhang, Bei Li, Ke Wang, Ben Xu
Phys. Rev. B, 2023, 107, 14101.
DOI: 10.1103/PhysRevB.107.014101
Li ion diffusion behavior of Li3OCl solid-state electrolytes with different defect structures: insights from the deep potential model
Zhou Zhang, Zhongyun Ma, Yong Pei
Phys. Chem. Chem. Phys., 2023, 25, 13297–13307.
DOI: 10.1039/d2cp06073f
A deep learning-based potential developed for calcium silicate hydrates with both high accuracy and efficiency
Weihuan Li, Yang Zhou, Li Ding, Pengfei Lv, Yifan Su, Rui Wang, Changwen Miao
Journal of Sustainable Cement-Based Materials, 2023, 12, 1335–1346.
DOI: 10.1080/21650373.2023.2219251
Tuning the lattice thermal conductivity of Sb2Te3 by Cr doping: a deep potential molecular dynamics study
Pan Zhang, Wenkai Liao, Ziyang Zhu, Mi Qin, Zhenhua Zhang, Dan Jin, Yong Liu, Ziyu Wang, Zhihong Lu, Rui Xiong
Phys. Chem. Chem. Phys., 2023, 25, 15422–15432.
DOI: 10.1039/d3cp00999h
A deep learning interatomic potential suitable for simulating radiation damage in bulk tungsten
Chang-Jie Ding, Ya-Wei Lei, Xiao-Yang Wang, Xiao-Lin Li, Xiang-Yan Li, Yan-Ge Zhang, Yi-Chun Xu, Chang-Song Liu, Xue-Bang Wu
Tungsten, 2023.
DOI: 10.1007/s42864-023-00230-4
Deep-learning potentials for proton transport in double-sided graphanol
Siddarth K. Achar, Leonardo Bernasconi, Juan J. Alvarez, J. Karl Johnson
Journal of Materials Research, 2023.
DOI: 10.1557/s43578-023-01141-3
A deep potential molecular dynamics study on the ionic structure and transport properties of NaCl-CaCl2 molten salt
Gegentana, Liu Cui, Leping Zhou, Xiaoze Du
Ionics, 2023, 1–11.
A first-principles machine-learning force field for heterogeneous ice nucleation on microcline feldspar
Pablo M Piaggi, Annabella Selloni, Athanassios Z Panagiotopoulos, Roberto Car, Pablo G Debenedetti
Faraday Discuss., 2023.
DOI: 10.1039/d3fd00100h
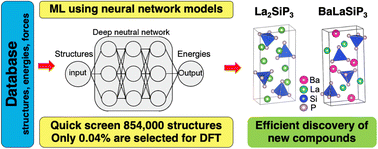
Accelerating materials discovery using integrated deep machine learning approaches
Weiyi Xia, Ling Tang, Huaijun Sun, Chao Zhang, Kai-Ming Ho, Gayatri Viswanathan, Kirill Kovnir, Cai-Zhuang Wang
J. Mater. Chem. A, 2023.
DOI: 10.1039/d3ta03771a
Speciation of La3+-Cl- Complexes in Hydrothermal Fluids from Deep Potential Molecular Dynamics
Wei Zhang, Li Zhou, Tinggui Yan, Mohan Chen
J. Phys. Chem. B, 2023, 127, 8926–8937.
DOI: 10.1021/acs.jpcb.3c05428
Neural Network Water Model Based on the MB-Pol Many-Body Potential
Maria Carolina Muniz, Roberto Car, Athanassios Z Panagiotopoulos
J. Phys. Chem. B, 2023, 127, 9165–9171.
DOI: 10.1021/acs.jpcb.3c04629
Understanding the Anomalous Diffusion of Water in Aqueous Electrolytes Using Machine Learned Potentials
Nikhil V S Avula, Michael L Klein, Sundaram Balasubramanian
J. Phys. Chem. Lett., 2023, 14, 9500–9507.
DOI: 10.1021/acs.jpclett.3c02112
Insights into the local structure evolution and thermophysical properties of NaCl-KCl-MgCl2\texte ndashLaCl3 melt driven by machine learning
Jia Zhao, Taixi Feng, Guimin Lu, Jianguo Yu
J. Mater. Chem. A, 2023, 11, 23999–24012.
DOI: 10.1039/d3ta03434h
Constrained Hybrid Monte Carlo Sampling Made Simple for Chemical Reaction Simulations
Bin Jin, Taiping Hu, Kuang Yu, Shenzhen Xu
J. Chem. Theory Comput., 2023, 19, 7343–7357.
DOI: 10.1021/acs.jctc.3c00571
Water dissociation at the water-rutile TiO 2 (110) interface from ab~initio-based deep neural network simulations
Bo Wen, Marcos F Calegari Andrade, Li-Min Liu, Annabella Selloni
Proc. Natl. Acad. Sci. U. S. A., 2023, 120, e2212250120.
DOI: 10.1073/pnas.2212250120
Development and Validation of Versatile Deep Atomistic Potentials for Metal Oxides
Pandu Wisesa, Christopher M Andolina, Wissam A Saidi
J. Phys. Chem. Lett., 2023, 14, 468–475.
DOI: 10.1021/acs.jpclett.2c03445
Machine learning assisted investigation of the barocaloric performance in ammonium iodide
Xiong Xu, Fangbiao Li, Chang Niu, Min Li, Hui Wang
2023, 122.
DOI: 10.1063/5.0131696
Accessing the thermal conductivities of Sb2Te3 and Bi2Te3/Sb2Te3 superlattices by molecular dynamics simulations with a deep neural network potential
Pan Zhang, Mi Qin, Zhenhua Zhang, Dan Jin, Yong Liu, Ziyu Wang, Zhihong Lu, Jing Shi, Rui Xiong
Phys. Chem. Chem. Phys., 2023, 25, 6164–6174.
DOI: 10.1039/d2cp05590b
Predicting Structural Properties of Pure Silica Zeolites Using Deep Neural Network Potentials
Tyler G Sours, Ambarish R Kulkarni
J. Phys. Chem. C. Nanomater. Interfaces, 2023, 127, 1455–1463.
DOI: 10.1021/acs.jpcc.2c08429
Melting of MgSiO3 determined by machine learning potentials
Jie Deng, Haiyang Niu, Junwei Hu, Mingyi Chen, Lars Stixrude
Phys. Rev. B, 2023, 107, 64103.
DOI: 10.1103/PhysRevB.107.064103
Machine learning based implicit solvent model for aqueous-solution alanine dipeptide molecular dynamics simulations
Songyuan Yao, Richard Van, Xiaoliang Pan, Ji Hwan Park, Yuezhi Mao, Jingzhi Pu, Ye Mei, Yihan Shao
RSC Adv., 2023, 13, 4565–4577.
DOI: 10.1039/d2ra08180f
Large-scale atomistic simulation of dislocation core structure in face-centered cubic metal with Deep Potential method
Fenglin Deng, Hongyu Wu, Ri He, Peijun Yang, Zhicheng Zhong
Computational Materials Science, 2023, 218, 111941.
DOI: 10.1016/j.commatsci.2022.111941
Melting conditions and entropies of superionic water ice: Free-energy calculations based on hybrid solid/liquid reference systems
Vitor Fidalgo C\^andido, Filipe Matusalem, Maurice de Koning
J. Chem. Phys., 2023, 158, 64502.
DOI: 10.1063/5.0138987
Size and Quality of Quantum Mechanical Data Set for Training Neural Network Force Fields for Liquid Water
M'arcio S Gomes-Filho, Alberto Torres, Alexandre Reily Rocha, Luana S Pedroza
J. Phys. Chem. B, 2023, 127, 1422–1428.
DOI: 10.1021/acs.jpcb.2c09059
Thermal transport across copper-water interfaces according to deep potential molecular dynamics
Zhiqiang Li, Xiaoyu Tan, Zhiwei Fu, Linhua Liu, Jia-Yue Yang
Phys. Chem. Chem. Phys., 2023, 25, 6746–6756.
DOI: 10.1039/d2cp05530a
Quasiclassical Trajectory Simulation as a Protocol to Build Locally Accurate Machine Learning Potentials
Jintu Zhang, Haotian Zhang, Zhixin Qin, Yu Kang, Xin Hong, Tingjun Hou
J. Chem. Inf. Model., 2023, 63, 1133–1142.
DOI: 10.1021/acs.jcim.2c01497
QD$\pi$: A Quantum Deep Potential Interaction Model for Drug Discovery
Jinzhe Zeng, Yujun Tao, Timothy J Giese, Darrin M York
J. Chem. Theory Comput., 2023, 19, 1261–1275.
DOI: 10.1021/acs.jctc.2c01172
A deep learning approach to predict thermophysical properties of metastable liquid Ti-Ni-Cr-Al alloy
R. L. Xiao, Q. Wang, J. Y. Qin, J. F. Zhao, Y. Ruan, H. P. Wang, H. Li, B. Wei
2023, 133.
DOI: 10.1063/5.0138001
A \textquotedblleftshort blanket\textquotedblright dilemma for a state-of-the-art neural network potential for water: Reproducing experimental properties or the physics of the underlying many-body interactions?
Yaoguang Zhai, Alessandro Caruso, Sigbj\orn L\oland Bore, Zhishang Luo, Francesco Paesani
J. Chem. Phys., 2023, 158, 84111.
DOI: 10.1063/5.0142843
Structure and solidification of the (Fe0.75B0.15Si0.1)100-xTax (x=0-2) melts: Experiment and machine learning
I.V. Sterkhova, L.V. Kamaeva, V.I. Lad'yanov, N.M. Chtchelkatchev
Journal of Physics and Chemistry of Solids, 2023, 174, 111143.
DOI: 10.1016/j.jpcs.2022.111143
Unravelling the dissolution dynamics of silicate minerals by deep learning molecular dynamics simulation: A case of dicalcium silicate
Yunjian Li, Hui Pan, Zongjin Li
Cement and Concrete Research, 2023, 165, 107092.
DOI: 10.1016/j.cemconres.2023.107092
Profiling the off-center atomic displacements in CuCl at finite temperatures with a deep-learning potential
Zhi-Hao Wang, Xuan-Yan Chen, Zhen Zhang, Xie Zhang, Su- Huai Wei
Phys. Rev. Materials, 2023, 7, 34601.
DOI: 10.1103/PhysRevMaterials.7.034601
Liquid-Crystal Structure Inheritance in Machine Learning Potentials for Network-Forming Systems
I. A. Balyakin, R. E. Ryltsev, N. M. Chtchelkatchev
Jetp Lett., 2023, 117, 370–376.
DOI: 10.1134/S0021364023600234
Oxygen Vacancy Diffusion in Rutile TiO2: Insight from Deep Neural Network Potential Simulations
Zhihong Wu, Wen-Jin Yin, Bo Wen, Dongwei Ma, Li-Min Liu
J. Phys. Chem. Lett., 2023, 14, 2208–2214.
DOI: 10.1021/acs.jpclett.2c03827
Nanotwinning-induced pseudoplastic deformation in boron carbide under low temperature
Jun Li, Qi An
International Journal of Mechanical Sciences, 2023, 242, 107998.
DOI: 10.1016/j.ijmecsci.2022.107998
Quantum Informed Machine-Learning Potentials for Molecular Dynamics Simulations of CO2’s Chemisorption and Diffusion in Mg-MOF-74
Bowen Zheng, Felipe Lopes Oliveira, Rodrigo Neumann Barros Ferreira, Mathias Steiner, Hendrik Hamann, Grace X Gu, Binquan Luan
ACS Nano, 2023, 17, 5579–5587.
DOI: 10.1021/acsnano.2c11102
Modern semiempirical electronic structure methods and machine learning potentials for drug discovery: Conformers, tautomers, and protonation states
Jinzhe Zeng, Yujun Tao, Timothy J Giese, Darrin M York
J. Chem. Phys., 2023, 158, 124110.
DOI: 10.1063/5.0139281
Innate dynamics and identity crisis of a metal surface unveiled by machine learning of atomic environments
Matteo Cioni, Daniela Polino, Daniele Rapetti, Luca Pesce, Massimo Delle Piane, Giovanni M Pavan
J. Chem. Phys., 2023, 158, 124701.
DOI: 10.1063/5.0139010
Prediction on local structure and properties of LiCl-KCl-AlCl3 ternary molten salt with deep learning potential
Min Bu, Taixi Feng, Guimin Lu
Journal of Molecular Liquids, 2023, 375, 120689.
DOI: 10.1016/j.molliq.2022.120689
Atomic structure, stability, and dissociation of dislocations in cadmium telluride
Jun Li, Kun Luo, Qi An
International Journal of Plasticity, 2023, 163, 103552.
DOI: 10.1016/j.ijplas.2023.103552
The highest melting point material: Searched by Bayesian global optimization with deep potential molecular dynamics
Yinan Wang, Bo Wen, Xingjian Jiao, Ya Li, Lei Chen, Yujin Wang, Fu-Zhi Dai
2023, 12, 803–814.
DOI: 10.26599/JAC.2023.9220721
High-Accuracy Neural Network Interatomic Potential for Silicon Nitride
Hui Xu, Zeyuan Li, Zhaofu Zhang, Sheng Liu, Shengnan Shen, Yuzheng Guo
Nanomaterials (Basel)., 2023, 13, 1352.
DOI: 10.3390/nano13081352
A New Spinel Chloride Solid Electrolyte with High Ionic Conductivity and Stability for Na-Ion Batteries
Jiahui Liu, Shuo Wang, Yoshiyuki Kawazoe, Qiang Sun
ACS Materials Lett., 2023, 5, 1009–1017.
DOI: 10.1021/acsmaterialslett.3c00119
Scalable hybrid deep neural networks/polarizable potentials biomolecular simulations including long-range effects
Th'eo Jaffrelot Inizan, Thomas Pl'e, Olivier Adjoua, Pengyu Ren, Hatice G"okcan, Olexandr Isayev, Louis Lagard\`ere, Jean-Philip Piquemal
Chem. Sci., 2023, 14, 5438–5452.
DOI: 10.1039/d2sc04815a
Structural and Composition Evolution of Palladium Catalyst for CO Oxidation under Steady-State Reaction Conditions
Jiawei Wu, Dingming Chen, Jianfu Chen, Haifeng Wang
J. Phys. Chem. C, 2023, 127, 6262–6270.
DOI: 10.1021/acs.jpcc.2c07877
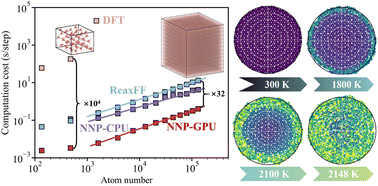
Monitoring the melting behavior of boron nanoparticles using a neural network potential
Xiaoya Chang, Qingzhao Chu, Dongping Chen
Phys. Chem. Chem. Phys., 2023, 25, 12841–12853.
DOI: 10.1039/d3cp00571b
Microstructure and Thermophysical Property Prediction for Chloride Composite Phase Change Materials: A Deep Potential Molecular Dynamics Study
C. Y. Zhao, Y. B. Tao, Y. He
J. Phys. Chem. C, 2023, 127, 6852–6860.
DOI: 10.1021/acs.jpcc.2c08589
Cooperative diffusion in body-centered cubic iron in Earth and super- Earths’ inner core conditions
Maitrayee Ghosh, Shuai Zhang, Lianming Hu, S X Hu
J. Phys. Condens. Matter, 2023, 35, 154002.
DOI: 10.1088/1361-648X/acba71
Pressure-Induced Stability of Methane Hydrate from Machine Learning Force Field Simulations
Kun Luo, Yidi Shen, Jun Li, Qi An
J. Phys. Chem. C, 2023, 127, 7071–7077.
DOI: 10.1021/acs.jpcc.2c09121
Unraveling the Dynamic Correlations between Transition Metal Migration and the Oxygen Dimer Formation in the Highly Delithiated LixCoO2 Cathode
Taiping Hu, Fu-Zhi Dai, Guobing Zhou, Xiaoxu Wang, Shenzhen Xu
J. Phys. Chem. Lett., 2023, 14, 3677–3684.
DOI: 10.1021/acs.jpclett.3c00506
Predicting thermodynamic stability of magnesium alloys in machine learning
Xi He, Jinde Liu, Chen Yang, Gang Jiang
Computational Materials Science, 2023, 223, 112111.
DOI: 10.1016/j.commatsci.2023.112111
Hydrogen distribution between the Earth's inner and outer core
Liang Yuan, Gerd Steinle-Neumann
Earth and Planetary Science Letters, 2023, 609, 118084.
DOI: 10.1016/j.epsl.2023.118084
Lattice Thermal Conductivity of Monolayer InSe Calculated by Machine Learning Potential
Jinsen Han, Qiyu Zeng, Ke Chen, Xiaoxiang Yu, Jiayu Dai
Nanomaterials (Basel)., 2023, 13, 1576.
DOI: 10.3390/nano13091576
Modeling the high-pressure solid and liquid phases of tin from deep potentials with ab initio accuracy
Tao Chen, Fengbo Yuan, Jianchuan Liu, Huayun Geng, Linfeng Zhang, Han Wang, Mohan Chen
Phys. Rev. Materials, 2023, 7, 53603.
DOI: 10.1103/PhysRevMaterials.7.053603
Estimation of frequency factors for the calculation of kinetic isotope effects from classical and path integral free energy simulations
Timothy J Giese, Darrin M York
J. Chem. Phys., 2023, 158.
DOI: 10.1063/5.0147218
A Deep Potential model for liquid-vapor equilibrium and cavitation rates of water
Ignacio Sanchez-Burgos, Maria Carolina Muniz, Jorge R Espinosa, Athanassios Z Panagiotopoulos
J. Chem. Phys., 2023, 158.
DOI: 10.1063/5.0144500
First-Principles-Based Machine Learning Models for Phase Behavior and Transport Properties of CO2
Reha Mathur, Maria Carolina Muniz, Shuwen Yue, Roberto Car, Athanassios Z Panagiotopoulos
J. Phys. Chem. B, 2023, 127, 4562–4569.
DOI: 10.1021/acs.jpcb.3c00610
Characterizing Structure-Dependent TiS2/Water Interfaces Using Deep-Neural-Network-Assisted Molecular Dynamics
Lesheng Li, Marcos F. Calegari Andrade, Roberto Car, Annabella Selloni, Emily A. Carter
J. Phys. Chem. C, 2023, 127, 9750–9758.
DOI: 10.1021/acs.jpcc.2c08581
An accurate interatomic potential for the TiAlNb ternary alloy developed by deep neural network learning method
Jiajun Lu, Jinkai Wang, Kaiwei Wan, Ying Chen, Hao Wang, Xinghua Shi
J. Chem. Phys., 2023, 158.
DOI: 10.1063/5.0147720
In Silico Demonstration of Fast Anhydrous Proton Conduction on Graphanol
Siddarth K Achar, Leonardo Bernasconi, Ruby I DeMaio, Katlyn R Howard, J Karl Johnson
ACS Appl. Mater. Interfaces, 2023, 15, 25873–25883.
DOI: 10.1021/acsami.3c04022
Machine learning insight into h-BN growth on Pt(111) from atomic states
Kangmo Yeo, Sukmin Jeong
Applied Surface Science, 2023, 621, 156893.
DOI: 10.1016/j.apsusc.2023.156893
Molecular dynamics simulations of CaCl2-NaCl molten salt based on the machine learning potentials
Yun Xie, Min Bu, Guiming Zou, Ye Zhang, Guimin Lu
Solar Energy Materials and Solar Cells, 2023, 254, 112275.
DOI: 10.1016/j.solmat.2023.112275
A Heterogeneous Parallel Non-von Neumann Architecture System for Accurate and Efficient Machine Learning Molecular Dynamics
Zhuoying Zhao, Ziling Tan, Pinghui Mo, Xiaonan Wang, Dan Zhao, Xin Zhang, Ming Tao, Jie Liu
IEEE Trans. Circuits Syst. I, 2023, 70, 2439–2449.
DOI: 10.1109/TCSI.2023.3255199
Noble gas (He, Ne, and Ar) solubilities in high-pressure silicate melts calculated based on deep-potential modeling
Kai Wang, Xiancai Lu, Xiandong Liu, Kun Yin
Geochimica et Cosmochimica Acta, 2023, 350, 57–68.
DOI: 10.1016/j.gca.2023.03.032
Reversible densification and cooperative atomic movement induced \textquotedblleftcompaction\textquotedblright in vitreous silica: a new sight from deep neural network interatomic potentials
Yongnian Qi, Xiaoguang Guo, Hao Wang, Shuohua Zhang, Ming Li, Ping Zhou, Dongming Guo
J Mater Sci, 2023, 58, 9515–9532.
DOI: 10.1007/s10853-023-08599-w
Molecular dynamics simulation of Fe-Si alloys using a neural network machine learning potential
Huaijun Sun, Chao Zhang, Ling Tang, Renhai Wang, Weiyi Xia, Cai-Zhuang Wang
Phys. Rev. B, 2023, 107, 224301.
DOI: 10.1103/PhysRevB.107.224301
Evaluation of Machine Learning Interatomic Potentials for Gold Nanoparticles\textemdashTransferability towards Bulk
Marco Fronzi, Roger D Amos, Rika Kobayashi
Nanomaterials (Basel)., 2023, 13, 1832.
DOI: 10.3390/nano13121832
A genome dependence of metastable phase selection on atomic structure for undercooled liquid Nb90Si10 hypoeutectic alloy
Q. Wang, C. H. Zheng, M. X. Li, L. Hu, H. P. Wang, B. Wei
2023, 122.
DOI: 10.1063/5.0152293
TimeSOAP: Tracking high-dimensional fluctuations in complex molecular systems via time variations of SOAP spectra
Cristina Caruso, Annalisa Cardellini, Martina Crippa, Daniele Rapetti, Giovanni M Pavan
J. Chem. Phys., 2023, 158.
DOI: 10.1063/5.0147025
Deciphering the Anomalous Acidic Tendency of Terminal Water at Rutile(110)-Water Interfaces
Yong-Bin Zhuang, Jun Cheng
J. Phys. Chem. C, 2023, 127, 10532–10540.
DOI: 10.1021/acs.jpcc.3c01870
Temperature-dependent microwave dielectric permittivity of gallium oxide: A deep potential molecular dynamics study
Zhiqiang Li, Xinlei Duan, Linhua Liu, Jia-Yue Yang
2023, 133.
DOI: 10.1063/5.0149447
Investigating the Hydroxyl Reorientation in Hydroxyapatite Using Machine Learning Potentials
Jing Wang, Xin Wang, Hua Zhu, Dingguo Xu
J. Phys. Chem. C, 2023, 127, 11369–11377.
DOI: 10.1021/acs.jpcc.3c02426
Probing Confinement Effects on the Infrared Spectra of Water with Deep Potential Molecular Dynamics Simulations
Marcos F Calegari Andrade, Tuan Anh Pham
J. Phys. Chem. Lett., 2023, 14, 5560–5566.
DOI: 10.1021/acs.jpclett.3c01054
A Deep Neural Network Potential to Study the Thermal Conductivity of MnBi2Te4 and Bi2Te3/MnBi2Te4 Superlattice
Ruijin Qu, Yawei Lv, Zhihong Lu
J. Electron. Mater., 2023, 52, 4475–4483.
DOI: 10.1007/s11664-023-10403-z
Molecular insight into the GaP(110)-water interface using machine learning accelerated molecular dynamics
Xue-Ting Fan, Xiao-Jian Wen, Yong-Bin Zhuang, Jun Cheng
Journal of Energy Chemistry, 2023, 82, 239–247.
DOI: 10.1016/j.jechem.2023.03.013
Modelling of dislocations, twins and crack-tips in HCP and BCC Ti
Tongqi Wen, Anwen Liu, Rui Wang, Linfeng Zhang, Jian Han, Han Wang, David J. Srolovitz, Zhaoxuan Wu
International Journal of Plasticity, 2023, 166, 103644.
DOI: 10.1016/j.ijplas.2023.103644
Halide Vacancies Create No Charge Traps on Lead Halide Perovskite Surfaces but Can Generate Deep Traps in the Bulk
Jingyi Ran, Bipeng Wang, Yifan Wu, Dongyu Liu, Carlos Mora Perez, Andrey S Vasenko, Oleg V Prezhdo
J. Phys. Chem. Lett., 2023, 14, 6028–6036.
DOI: 10.1021/acs.jpclett.3c01231
High-Throughput Condensed-Phase Hybrid Density Functional Theory for Large-Scale Finite-Gap Systems: The SeA Approach
Hsin-Yu Ko, Marcos F Calegari Andrade, Zachary M Sparrow, Ju-An Zhang, Robert A DiStasio Jr
J. Chem. Theory Comput., 2023, 19, 4182–4201.
DOI: 10.1021/acs.jctc.2c00827
Microscopic Mechanism of Proton Transfer in Pure Water under Ambient Conditions
Jun Huo, Jianghao Chen, Pei Liu, Benkun Hong, Jian Zhang, Hao Dong, Shuhua Li
J. Chem. Theory Comput., 2023, 19, 4243–4254.
DOI: 10.1021/acs.jctc.3c00244
Revealing Carbon Vacancy Distribution on $\alpha$-MoC1-x Surfaces by Machine-Learning Force-Field-Aided Cluster Expansion Approach
Jun-Zhong Xie, Hong Jiang
J. Phys. Chem. C, 2023, 127, 13228–13237.
DOI: 10.1021/acs.jpcc.3c01941
Modeling the aqueous interface of amorphous TiO2 using deep potential molecular dynamics
Zhutian Ding, Annabella Selloni
J. Chem. Phys., 2023, 159.
DOI: 10.1063/5.0157188
Transferability evaluation of the deep potential model for simulating water-graphene confined system
Dongfei Liu, Jianzhong Wu, Diannan Lu
J. Chem. Phys., 2023, 159.
DOI: 10.1063/5.0153196
Structural transformations in single-crystalline AgPd nanoalloys from multiscale deep potential molecular dynamics
Longfei Guo, Tao Jin, Shuang Shan, Quan Tang, Zhen Li, Chongyang Wang, Junpeng Wang, Bowei Pan, Qiao Wang, Fuyi Chen
J. Chem. Phys., 2023, 159.
DOI: 10.1063/5.0158918
Detecting dynamic domains and local fluctuations in complex molecular systems via timelapse neighbors shuffling
Martina Crippa, Annalisa Cardellini, Cristina Caruso, Giovanni M Pavan
Proc. Natl. Acad. Sci. U. S. A., 2023, 120, e2300565120.
DOI: 10.1073/pnas.2300565120
Anisotropic Collective Variables with Machine Learning Potential for Ab Initio Crystallization of Complex Ceramics
Yuanpeng Deng, Shubin Fu, Jingran Guo, Xiang Xu, Hui Li
ACS Nano, 2023, 17, 14099–14113.
DOI: 10.1021/acsnano.3c04602
Fluorine spillover for ceria- vs silica-supported palladium nanoparticles: A MD study using machine learning potentials
Da-Jiang Liu, James W Evans
J. Chem. Phys., 2023, 159.
DOI: 10.1063/5.0147132
Rapid acquisition of liquid thermophysical properties from pure metals to quaternary alloys by proposing a machine learning strategy
R. L. Xiao, K. L. Liu, Y. Ruan, B. Wei
2023, 123.
DOI: 10.1063/5.0160046
Extreme phonon anharmonicity underpins superionic diffusion and ultralow thermal conductivity in argyrodite Ag8SnSe6
Qingyong Ren, Mayanak K Gupta, Min Jin, Jingxuan Ding, Jiangtao Wu, Zhiwei Chen, Siqi Lin, Oscar Fabelo, Jose Alberto Rodr'\iguez-Velamaz'an, Maiko Kofu, Kenji Nakajima, Marcell Wolf, Fengfeng Zhu, Jianli Wang, Zhenxiang Cheng, Guohua Wang, Xin Tong, Yanzhong Pei, Olivier Delaire, Jie Ma
Nat. Mater., 2023, 22, 999–1006.
DOI: 10.1038/s41563-023-01560-x
Highly transferable atomistic machine-learning potentials from curated and compact datasets across the periodic table
Christopher M. Andolina, Wissam A. Saidi
Digital Discovery, 2023, 2, 1070–1077.
DOI: 10.1039/d3dd00046j
Unraveling the Oxidation Behaviors of MXenes in Aqueous Systems by Active-Learning-Potential Molecular-Dynamics Simulation
Pengfei Hou, Yumiao Tian, Yu Xie, Fei Du, Gang Chen, Aleksandra Vojvodic, Jianzhong Wu, Xing Meng
Angew. Chem. Int. Ed. Engl., 2023, 62, e202304205.
DOI: 10.1002/anie.202304205
DeePMD-kit v2: A software package for deep potential models
Jinzhe Zeng, Duo Zhang, Denghui Lu, Pinghui Mo, Zeyu Li, Yixiao Chen, Mari'an Rynik, Li'ang Huang, Ziyao Li, Shaochen Shi, Yingze Wang, Haotian Ye, Ping Tuo, Jiabin Yang, Ye Ding, Yifan Li, Davide Tisi, Qiyu Zeng, Han Bao, Yu Xia, Jiameng Huang, Koki Muraoka, Yibo Wang, Junhan Chang, Fengbo Yuan, Sigbj\orn L\oland Bore, Chun Cai, Yinnian Lin, Bo Wang, Jiayan Xu, Jia-Xin Zhu, Chenxing Luo, Yuzhi Zhang, Rhys E A Goodall, Wenshuo Liang, Anurag Kumar Singh, Sikai Yao, Jingchao Zhang, Renata Wentzcovitch, Jiequn Han, Jie Liu, Weile Jia, Darrin M York, Weinan E, Roberto Car, Linfeng Zhang, Han Wang
J. Chem. Phys., 2023, 159.
DOI: 10.1063/5.0155600
Melting curves of ice polymorphs in the vicinity of the liquid-liquid critical point
Pablo M Piaggi, Thomas E Gartner, Roberto Car, Pablo G Debenedetti
J. Chem. Phys., 2023, 159.
DOI: 10.1063/5.0159288
Spontaneous Hybrid Nano-Domain Behavior of the Organic-Inorganic Hybrid Perovskites
Ping Tuo, Lei Li, Xiaoxu Wang, Jianhui Chen, Zhicheng Zhong, Bo Xu, Fu-Zhi Dai
Adv. Funct. Mater., 2023, 33.
DOI: 10.1002/adfm.202301663
Thermal Conductivity of Water at Extreme Conditions
Cunzhi Zhang, Marcello Puligheddu, Linfeng Zhang, Roberto Car, Giulia Galli
J. Phys. Chem. B, 2023, 127, 7011–7.
DOI: 10.1021/acs.jpcb.3c02972
Local structure, thermodynamics, and melting of boron phosphide at high pressures by deep learning-driven ab~initio simulations
N M Chtchelkatchev, R E Ryltsev, M V Magnitskaya, S M Gorbunov, K A Cherednichenko, V L Solozhenko, V V Brazhkin
J. Chem. Phys., 2023, 159.
DOI: 10.1063/5.0165948
Exploring Configurations of Nanocrystal Ligands Using Machine-Learned Force Fields
Jakub K Sowa, Sean T Roberts, Peter J Rossky
J. Phys. Chem. Lett., 2023, 14, 7215–7222.
DOI: 10.1021/acs.jpclett.3c01618
Deep Potential Molecular Dynamics Study of Chapman-Jouguet Detonation Events of Energetic Materials
Jidong Zhang, Wei Guo, Yugui Yao
J. Phys. Chem. Lett., 2023, 14, 7141–7148.
DOI: 10.1021/acs.jpclett.3c01392
Why Dissolving Salt in Water Decreases Its Dielectric Permittivity
Chunyi Zhang, Shuwen Yue, Athanassios Z Panagiotopoulos, Michael L Klein, Xifan Wu
Phys. Rev. Lett., 2023, 131, 76801.
DOI: 10.1103/PhysRevLett.131.076801
Characterization of the Hydrogen-Bond Network in High-Pressure Water by Deep Potential Molecular Dynamics
Renxi Liu, Mohan Chen
J. Chem. Theory Comput., 2023, 19, 5602–5608.
DOI: 10.1021/acs.jctc.3c00445
Equation of state predictions for ScF3 and CaZrF6 with neural network- driven molecular dynamics
John P Stoppelman, Angus P Wilkinson, Jesse G McDaniel
J. Chem. Phys., 2023, 159.
DOI: 10.1063/5.0157615
Reactant-induced dynamics of lithium imide surfaces during the ammonia decomposition process
Manyi Yang, Umberto Raucci, Michele Parrinello
Nat Catal, 2023, 6, 829–836.
DOI: 10.1038/s41929-023-01006-2
Deep neural network potential for simulating hydrogen blistering in tungsten
Xiao-Yang Wang, Yi-Nan Wang, Ke Xu, Fu-Zhi Dai, Hai-Feng Liu, Guang-Hong Lu, Han Wang
Phys. Rev. Materials, 2023, 7, 93601.
DOI: 10.1103/PhysRevMaterials.7.093601
ChecMatE: A workflow package to automatically generate machine learning potentials and phase diagrams for semiconductor alloys
Yu-Xin Guo, Yong-Bin Zhuang, Jueli Shi, Jun Cheng
J. Chem. Phys., 2023, 159.
DOI: 10.1063/5.0166858
Unraveling the Atomic-scale Mechanism of Phase Transformations and Structural Evolutions during (de)Lithiation in Si Anodes
Fangjia Fu, Xiaoxu Wang, Linfeng Zhang, Yifang Yang, Jianhui Chen, Bo Xu, Chuying Ouyang, Shenzhen Xu, Fu-Zhi Dai, Weinan E
Adv. Funct. Mater., 2023, 33.
DOI: 10.1002/adfm.202303936
High-Accuracy Machine-Learned Interatomic Potentials for the Phase Change Material Ge3Sb6Te5
Wei Yu, Zhaofu Zhang, Xuhao Wan, Jinhao Su, Qingzhong Gui, Hailing Guo, Hong-xia Zhong, John Robertson, Yuzheng Guo
Chem. Mater., 2023, 35, 6651–6658.
DOI: 10.1021/acs.chemmater.3c00524

Distinct anharmonic characteristics of phonon-driven lattice thermal conductivity and thermal expansion in bulk MoSe2 and WSe2
Mayanak K. Gupta, Sajan Kumar, Ranjan Mittal, Sanjay K. Mishra, Stephane Rols, Olivier Delaire, Arumugum Thamizhavel, P. U. Sastry, Samrath L. Chaplot
J. Mater. Chem. A, 2023, 11, 21864–21873.
DOI: 10.1039/d3ta03830k
Deciphering the Atomistic Mechanism of Si(111)-7 \texttimes 7 Surface Reconstruction Using a Machine-Learning Force Field
Yidi Shen, Sergey I Morozov, Kun Luo, Qi An, William A Goddard Iii
J. Am. Chem. Soc., 2023, 145, 20511–20520.
DOI: 10.1021/jacs.3c06540
Large scale and quantum accurate molecular dynamics simulation: Liquid iron under extreme condition
Qi-Yu Zeng, Bo Chen, Dong-Dong Kang, Jia-Yu Dai
Acta Phys. Sin., 2023, 72, 187102.
DOI: 10.7498/aps.72.20231258
Role of hydrogen-doping for compensating oxygen-defect in non- stoichiometric amorphous In2O3$-$x: Modeling with a machine-learning potential
Shingo Urata, Nobuhiro Nakamura, Junghwan Kim, Hideo Hosono
2023, 134.
DOI: 10.1063/5.0149199
Advancing Accurate and Efficient Surface Behavior Modeling of Al Clusters with Machine Learning Potential
Chongteng Wu, Tong Liu, Xiayu Ran, Yuefeng Su, Yun Lu, Ning Li, Lai Chen, Zhenwei Wu, Feng Wu, Duanyun Cao
J. Phys. Chem. C, 2023, 127, 19115–19126.
DOI: 10.1021/acs.jpcc.3c03229
Atomistic modeling of lithium materials from deep learning potential with ab initio accuracy
Haidi Wang, Tao Li, Yufan Yao, Xiaofeng Liu, Weiduo Zhu, Zhao Chen, Zhongjun Li, Wei Hu
2023, 36, 573–581.
DOI: 10.1063/1674-0068/cjcp2211173
Collective motion in hcp-Fe at Earth’s inner core conditions
Youjun Zhang, Yong Wang, Yuqian Huang, Junjie Wang, Zhixin Liang, Long Hao, Zhipeng Gao, Jun Li, Qiang Wu, Hong Zhang, Yun Liu, Jian Sun, Jung-Fu Lin
Proc. Natl. Acad. Sci. U. S. A., 2023, 120, e2309952120.
DOI: 10.1073/pnas.2309952120
Machine-Learning Accelerated First-Principles Accurate Modeling of the Solid-Liquid Phase Transition in MgO under Mantle Conditions
Pandu Wisesa, Christopher M Andolina, Wissam A Saidi
J. Phys. Chem. Lett., 2023, 14, 8741–8748.
DOI: 10.1021/acs.jpclett.3c02424
Thermal transport across TiO2-H2O interface involving water dissociation: Ab initio-assisted deep potential molecular dynamics
Zhiqiang Li, Jian Wang, Chao Yang, Linhua Liu, Jia-Yue Yang
J. Chem. Phys., 2023, 159.
DOI: 10.1063/5.0167238
Deep-learning-assisted theoretical insights into the compatibility of environment friendly insulation medium with metal surface of power equipment
Xuhao Wan, Zhaofu Zhang, Anyang Wang, Jinhao Su, Wenjun Zhou, John Robertson, Yuan Peng, Yu Zheng, Yuzheng Guo
J. Colloid Interface Sci., 2023, 648, 317–326.
DOI: 10.1016/j.jcis.2023.05.188
Computational study of Na diffusion and conduction in P2- and O3-Na2x[NixTi1-x]O2 materials with machine-learning interatomic potentials
Yining He, Qian Chen, Wei Lai
Solid State Ionics, 2023, 399, 116298.
DOI: 10.1016/j.ssi.2023.116298
Self-powered H2 generation implemented by hydrazine oxidation assisting hybrid electrochemical cell
Xi Liu, Wei Sun, Xiang Hu, Junxiang Chen, Zhenhai Wen
Chemical Engineering Journal, 2023, 474, 145355.
DOI: 10.1016/j.cej.2023.145355
Understanding the sluggish and highly variable transport kinetics of lithium ions in LiFePO4
Youcheng Hu, Xiaoxiao Wang, Peng Li, Junxiang Chen, Shengli Chen
Sci. China Chem., 2023, 66, 3297–3306.
DOI: 10.1007/s11426-023-1662-9
Thermal conductivity of Fe-bearing bridgmanite and post-perovskite: Implications for the heat flux from the core
Dong Wang, Zhongqing Wu, Xin Deng
Earth and Planetary Science Letters, 2023, 621, 118368.
DOI: 10.1016/j.epsl.2023.118368
Inorganic glass electrolytes with polymer-like viscoelasticity
Tao Dai, Siyuan Wu, Yaxiang Lu, Yang Yang, Yuan Liu, Chao Chang, Xiaohui Rong, Ruijuan Xiao, Junmei Zhao, Yanhui Liu, Weihua Wang, Liquan Chen, Yong-Sheng Hu
Nat Energy, 2023, 8, 1221–1228.
DOI: 10.1038/s41560-023-01356-y
Machine learning potential for Ab Initio phase transitions of zirconia
Yuanpeng Deng, Chong Wang, Xiang Xu, Hui Li
Theoretical and Applied Mechanics Letters, 2023, 13, 100481.
DOI: 10.1016/j.taml.2023.100481
Modelling electrified microporous carbon/electrolyte electrochemical interface and unraveling charge storage mechanism by machine learning accelerated molecular dynamics
Yifeng Zhang, Hui Huang, Jie Tian, Chengwei Li, Yuchen Jiang, Zeng Fan, Lujun Pan
Energy Storage Materials, 2023, 63, 103069.
DOI: 10.1016/j.ensm.2023.103069
From ab~initio to continuum: Linking multiple scales using deep-learned forces
Haiyi Wu, Chenxing Liang, Jinu Jeong, N R Aluru
J. Chem. Phys., 2023, 159.
DOI: 10.1063/5.0166927
Superionic effect and anisotropic texture in Earth’s inner core driven by geomagnetic field
Shichuan Sun, Yu He, Junyi Yang, Yufeng Lin, Jinfeng Li, Duck Young Kim, Heping Li, Ho-Kwang Mao
Nat. Commun., 2023, 14, 1656.
DOI: 10.1038/s41467-023-37376-1
Data-driven prediction of complex crystal structures of dense lithium
Xiaoyang Wang, Zhenyu Wang, Pengyue Gao, Chengqian Zhang, Jian Lv, Han Wang, Haifeng Liu, Yanchao Wang, Yanming Ma
Nat. Commun., 2023, 14, 2924.
DOI: 10.1038/s41467-023-38650-y
Realistic phase diagram of water from \textquotedblleftfirst principles\textquotedblright data-driven quantum simulations
Sigbj\orn L\oland Bore, Francesco Paesani
Nat. Commun., 2023, 14, 3349.
DOI: 10.1038/s41467-023-38855-1
Realizing long-cycling all-solid-state Li-In||TiS2 batteries using Li6+xMxAs1-xS5I (M=Si, Sn) sulfide solid electrolytes
Pushun Lu, Yu Xia, Guochen Sun, Dengxu Wu, Siyuan Wu, Wenlin Yan, Xiang Zhu, Jiaze Lu, Quanhai Niu, Shaochen Shi, Zhengju Sha, Liquan Chen, Hong Li, Fan Wu
Nat. Commun., 2023, 14, 4077.
DOI: 10.1038/s41467-023-39686-w
Temperature-pressure phase diagram of confined monolayer water/ice at first-principles accuracy with a machine-learning force field
Bo Lin, Jian Jiang, Xiao Cheng Zeng, Lei Li
Nat. Commun., 2023, 14, 4110.
DOI: 10.1038/s41467-023-39829-z
Discrepancies and error evaluation metrics for machine learning interatomic potentials
Yunsheng Liu, Xingfeng He, Yifei Mo
npj Comput. Mater., 2023, 9, 174.
DOI: 10.1038/s41524-023-01123-3
Dislocation-mediated migration of the $\alpha$/$\beta$ interfaces in titanium
Jin-Yu Zhang, Zhi-Peng Sun, Dong Qiu, Fu-Zhi Dai, Yang- Sheng Zhang, Dongsheng Xu, Wen-Zheng Zhang
Acta Materialia, 2023, 261, 119364.
DOI: 10.1016/j.actamat.2023.119364
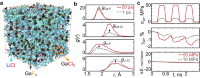
What dictates soft clay-like lithium superionic conductor formation from rigid salts mixture
Sunny Gupta, Xiaochen Yang, Gerbrand Ceder
Nat. Commun., 2023, 14, 6884.
DOI: 10.1038/s41467-023-42538-2
Interfacial heat and mass transfer at silica/binary molten salt interface from deep potential molecular dynamics
Fei Liang, Jing Ding, Xiaolan Wei, Gechuanqi Pan, Shule Liu
International Journal of Heat and Mass Transfer, 2023, 217, 124705.
DOI: 10.1016/j.ijheatmasstransfer.2023.124705
Reversible and irreversible photon-absorption in amorphous SiO2 revealed by deep potential
Yongnian Qi, Xiaoguang Guo, Ming Li, Chongkun Wang, Qing Mu, Ping Zhou
Journal of Non-Crystalline Solids, 2023, 622, 122682.
DOI: 10.1016/j.jnoncrysol.2023.122682
Thermal conductivity of van der Waals heterostructure of 2D GeS and SnS based on machine learning interatomic potential
Wentao Li, Chenxiu Yang
J. Phys. Condens. Matter, 2023, 35, 505001.
DOI: 10.1088/1361-648X/acf6ea
Development of Deep Potentials of Molten MgCl2-NaCl and MgCl2-KCl Salts Driven by Machine Learning
Tingrui Xu, Xuejiao Li, Yang Wang, Zhongfeng Tang
ACS Appl. Mater. Interfaces, 2023.
DOI: 10.1021/acsami.2c19272
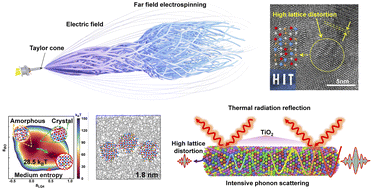
Medium-entropy ceramic aerogels for robust thermal sealing
Shubin Fu, Dizhou Liu, Yuanpeng Deng, Menglin Li, Han Zhao, Jingran Guo, Jian Zhou, Pengyu Zhang, Chong Wang, Hongxuan Yu, Shixuan Dang, Jianing Zhang, Menglong Hao, Hui Li, Xiang Xu
J. Mater. Chem. A, 2023, 11, 742–752.
DOI: 10.1039/d2ta08264k
Grain boundaries induce significant decrease in lattice thermal conductivity of CdTe
Xiaona Huang, Kun Luo, Yidi Shen, Yanan Yue, Qi An
Energy and AI, 2023, 11, 100210.
DOI: 10.1016/j.egyai.2022.100210
Revisiting the structure, interaction, and dynamical property of ionic liquid from the deep learning force field
Yulong Ling, Kun Li, Mi Wang, Junfeng Lu, Chenlu Wang, Yanlei Wang, Hongyan He
Journal of Power Sources, 2023, 555, 232350.
DOI: 10.1016/j.jpowsour.2022.232350
Quasiplastic deformation in shocked nanocrystalline boron carbide: Grain boundary sliding and local amorphization
Jun Li, Qi An
Journal of the European Ceramic Society, 2023, 43, 208–216.
DOI: 10.1016/j.jeurceramsoc.2022.10.014
Accurate Fe-He machine learning potential for studying He effects in BCC-Fe
Krishna Chaitanya Pitike, Wahyu Setyawan
Journal of Nuclear Materials, 2023, 574, 154183.
DOI: 10.1016/j.jnucmat.2022.154183
Solvation structures of calcium and magnesium ions in water with the presence of hydroxide: a study by deep potential molecular dynamics
Jianchuan Liu, Renxi Liu, Yu Cao, Mohan Chen
Phys. Chem. Chem. Phys., 2023.
DOI: 10.1039/d2cp04105g
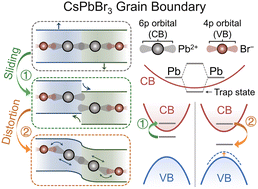
Grain boundary sliding and distortion on a nanosecond timescale induce trap states in CsPbBr3: ab initio investigation with machine learning force field
Dongyu Liu, Yifan Wu, Andrey S Vasenko, Oleg V Prezhdo
Nanoscale, 2023, 15, 285–293.
DOI: 10.1039/d2nr05918e
Accurate interatomic potential for the nucleation in liquid Ti-Al binary alloy developed by deep neural network learning method
B. Zhai, H.P. Wang
Computational Materials Science, 2023, 216, 111843.
DOI: 10.1016/j.commatsci.2022.111843
Learning DeePMD-Kit: A Guide to Building Deep Potential Models
Wenshuo Liang, Jinzhe Zeng, Darrin M. York, Linfeng Zhang, Han Wang
A Practical Guide to Recent Advances in Multiscale Modeling and Simulation of Biomolecules, 2023, 6-1–6-20.
DOI: 10.1063/9780735425279_006
2022
Convergence acceleration in machine learning potentials for atomistic simulations
Dylan Bayerl, Christopher M. Andolina, Shyam Dwaraknath, Wissam A. Saidi
Digital Discovery, 2022, 1, 61–69.
DOI: 10.1039/d1dd00005e
Towards fully ab initio simulation of atmospheric aerosol nucleation
Shuai Jiang, Yi-Rong Liu, Teng Huang, Ya-Juan Feng, Chun- Yu Wang, Zhong-Quan Wang, Bin-Jing Ge, Quan-Sheng Liu, Wei-Ran Guang, Wei Huang
Nat. Commun., 2022, 13, 6067.
DOI: 10.1038/s41467-022-33783-y
Thermoelectric Figure of Merit of a Superatomic Crystal ReSeI2 Monolayer
Tingwei Li, Peng-Hu Du, Ling Bai, Qiang Sun, Puru Jena
Phys. Rev. Applied, 2022, 18, 64067.
DOI: 10.1103/PhysRevApplied.18.064067
Deep learning molecular dynamics simulation on microwave high- temperature dielectric function of silicon nitride
Zhi-Qiang Li, Xiao-Yu Tan, Xin-Lei Duan, Jing-Yi Zhang, Jia-Yue Yang
Acta Phys. Sin., 2022, 71, 247803.
DOI: 10.7498/aps.71.20221002
Transferable Deep Learning Potential Reveals Intermediate-Range Ordering Effects in LiF-NaF-ZrF4 Molten Salt
Rajni Chahal, Santanu Roy, Martin Brehm, Shubhojit Banerjee, Vyacheslav Bryantsev, Stephen T Lam
JACS Au, 2022, 2, 2693–2702.
DOI: 10.1021/jacsau.2c00526
Elasticity and Viscosity of hcp Iron at Earth's Inner Core Conditions From Machine Learning-Based Large-Scale Atomistic Simulations
Zhi Li, Sandro Scandolo
Geophysical Research Letters, 2022, 49.
DOI: 10.1029/2022GL101161
Modeling Chemical Reactions in Alkali Carbonate-Hydroxide Electrolytes with Deep Learning Potentials
Anirban Mondal, Dina Kussainova, Shuwen Yue, Athanassios Z Panagiotopoulos
J. Chem. Theory Comput., 2022.
DOI: 10.1021/acs.jctc.2c00816
Spatial arrangement of dynamic surface species from solid-state NMR and machine learning-accelerated MD simulations
Takeshi Kobayashi, Da-Jiang Liu, Fr'ed'eric A Perras
Chem. Commun. (Camb)., 2022, 58, 13939–13942.
DOI: 10.1039/d2cc05861h
Fluctuations at Metal Halide Perovskite Grain Boundaries Create Transient Trap States: Machine Learning Assisted Ab Initio Analysis
Yifan Wu, Dongyu Liu, Weibin Chu, Bipeng Wang, Andrey S Vasenko, Oleg V Prezhdo
ACS Appl. Mater. Interfaces, 2022, 14, 55753–55761.
DOI: 10.1021/acsami.2c16203
Modeling Short-Range and Three-Membered Ring Structures in Lithium Borosilicate Glasses Using a Machine-Learning Potential
Shingo Urata
J. Phys. Chem. C, 2022, 126, 21507–21517.
DOI: 10.1021/acs.jpcc.2c07597
Lattice Thermal Conductivity of MgSiO3 Perovskite and Post- Perovskite under Lower Mantle Conditions Calculated by Deep Potential Molecular Dynamics
Fenghu Yang, Qiyu Zeng, Bo Chen, Dongdong Kang, Shen Zhang, Jianhua Wu, Xiaoxiang Yu, Jiayu Dai
Chinese Phys. Lett., 2022, 39, 116301.
DOI: 10.1088/0256-307X/39/11/116301
Origin of the herringbone reconstruction of Au(111) surface at the atomic scale
Pai Li, Feng Ding
Sci. Adv., 2022, 8, eabq2900.
DOI: 10.1126/sciadv.abq2900
Resolving the odd-even oscillation of water dissociation at rutile TiO2(110)-water interface by machine learning accelerated molecular dynamics
Yong-Bin Zhuang, Rui-Hao Bi, Jun Cheng
J. Chem. Phys., 2022, 157, 164701.
DOI: 10.1063/5.0126333
Evaluation of Machine Learning Interatomic Potentials for the Properties of Gold Nanoparticles
Marco Fronzi, Roger D Amos, Rika Kobayashi, Naoki Matsumura, Kenta Watanabe, Rafael K Morizawa
Nanomaterials (Basel)., 2022, 12, 3891.
DOI: 10.3390/nano12213891
Predicted superconductivity and superionic state in the electride Li5N under high pressure
Zhongyu Wan, Chao Zhang, Tianyi Yang, Wenjun Xu, Ruiqin Zhang
New J. Phys., 2022, 24, 113012.
DOI: 10.1088/1367-2630/ac9cff
Origin of negative thermal expansion and pressure-induced amorphization in zirconium tungstate from a machine-learning potential
Ri He, Hongyu Wu, Yi Lu, Zhicheng Zhong
Phys. Rev. B, 2022, 106, 174101.
DOI: 10.1103/PhysRevB.106.174101
Phonon Thermal Transport in Bi2Te3 from a Deep-Neural-Network Interatomic Potential
Robert D McMichael, Sean M Blakley
Phys. Rev. Appl., 2022, 18, 54022.
DOI: 10.1103/PhysRevApplied.18.054022
Piezo- and Pyroelectricity in Zirconia: A Study with Machine-Learned Force Fields
Robert D McMichael, Sean M Blakley
Phys. Rev. Appl., 2022, 18, 54066.
DOI: 10.1103/PhysRevApplied.18.054066
Classical and machine learning interatomic potentials for BCC vanadium
Rui Wang, Xiaoxiao Ma, Linfeng Zhang, Han Wang, David J. Srolovitz, Tongqi Wen, Zhaoxuan Wu
Phys. Rev. Materials, 2022, 6, 113603.
DOI: 10.1103/PhysRevMaterials.6.113603
Order of magnitude reduction in Joule heating of single molecular junctions between graphene electrodes
Gen Li, Bing-Zhong Hu, Wen-Hao Mao, Nuo Yang, Jing-Tao L"u
J. Chem. Phys., 2022, 157, 174303.
DOI: 10.1063/5.0118952
Plastic deformation of superionic water ices
Filipe Matusalem, J'essica Santos Rego, Maurice de Koning
Proc. Natl. Acad. Sci. U. S. A., 2022, 119, e2203397119.
DOI: 10.1073/pnas.2203397119
Metal Affinity of Support Dictates Sintering of Gold Catalysts
Jin-Cheng Liu, Langli Luo, Hai Xiao, Junfa Zhu, Yang He, Jun Li
J. Am. Chem. Soc., 2022, 144, 20601–20609.
DOI: 10.1021/jacs.2c06785
Multireference Generalization of the Weighted Thermodynamic Perturbation Method
Timothy J Giese, Jinzhe Zeng, Darrin M York
J. Phys. Chem. A, 2022, 126, 8519–8533.
DOI: 10.1021/acs.jpca.2c06201
Thermal Conductivity of Hydrous Wadsleyite Determined by Non-Equilibrium Molecular Dynamics Based on Machine Learning
Dong Wang, Zhongqing Wu, Xin Deng
Geophysical Research Letters, 2022, 49.
DOI: 10.1029/2022GL100337
DeePKS + ABACUS as a Bridge between Expensive Quantum Mechanical Models and Machine Learning Potentials
Wenfei Li, Qi Ou, Yixiao Chen, Yu Cao, Renxi Liu, Chunyi Zhang, Daye Zheng, Chun Cai, Xifan Wu, Han Wang, Mohan Chen, Linfeng Zhang
J. Phys. Chem. A, 2022, 126, 9154–9164.
DOI: 10.1021/acs.jpca.2c05000
Centroid Molecular Dynamics Can Be Greatly Accelerated through Neural Network Learned Centroid Forces Derived from Path Integral Molecular Dynamics
Timothy D Loose, Patrick G Sahrmann, Gregory A Voth
J. Chem. Theory Comput., 2022, 18, 5856–5863.
DOI: 10.1021/acs.jctc.2c00706
Moir'e Phonons in Magic-Angle Twisted Bilayer Graphene
Xiaoqian Liu, Ran Peng, Zhaoru Sun, Jianpeng Liu
Nano Lett., 2022, 22, 7791–7797.
DOI: 10.1021/acs.nanolett.2c02010
Modeling the Solvation and Acidity of Carboxylic Acids Using an Ab Initio Deep Neural Network Potential
Abhinav S Raman, Annabella Selloni
J. Phys. Chem. A, 2022, 126, 7283–7290.
DOI: 10.1021/acs.jpca.2c06252
Photoelectron spectra of water and simple aqueous solutions at extreme conditions
Zifan Ye, Cunzhi Zhang, Giulia Galli
Faraday Discuss., 2022, 236, 352–363.
DOI: 10.1039/d2fd00003b
Deep potential for a face-centered cubic Cu system at finite temperatures
Yunzhen Du, Zhaocang Meng, Qiang Yan, Canglong Wang, Yuan Tian, Wenshan Duan, Sheng Zhang, Ping Lin
Phys. Chem. Chem. Phys., 2022, 24, 18361–18369.
DOI: 10.1039/D2CP02758E
Thermal transport properties of monolayer GeS and SnS: A comparative study based on machine learning and SW interatomic potential models
Wentao Li, Chenxiu Yang
AIP Advances, 2022, 12, 85111.
DOI: 10.1063/5.0099448
Structural and electrocatalytic properties of copper clusters: A study via deep learning and first principles
Xiaoning Wang, Haidi Wang, Qiquan Luo, Jinlong Yang
J. Chem. Phys., 2022, 157, 74304.
DOI: 10.1063/5.0100505
A Deep Neural Network Interface Potential for Li-Cu Systems
Genming Lai, Junyu Jiao, Chi Fang, Ruiqi Zhang, Xianqi Xu, Liyuan Sheng, Yao Jiang, Chuying Ouyang, Jiaxin Zheng
Adv. Mater. Interfaces, 2022, 9, 2201346.
DOI: 10.1002/admi.202201346
Strategy to consider element distribution when constructing training datasets for developing machine learning potentials of alloys based on a Monte-Carlo-like method
Zhipeng Zhang, Liuqing Chen, Junyi Guo, Xianyin Duan, Bin Shan, Xianbao Duan
Phys. Rev. B, 2022, 106, 94107.
DOI: 10.1103/PhysRevB.106.094107
Combining Machine Learning Approaches and Accurate Ab Initio Enhanced Sampling Methods for Prebiotic Chemical Reactions in Solution
Timoth'ee Devergne, Th'eo Magrino, Fabio Pietrucci, A Marco Saitta
J. Chem. Theory Comput., 2022, 18, 5410–5421.
DOI: 10.1021/acs.jctc.2c00400
GPUMD: A package for constructing accurate machine-learned potentials and performing highly efficient atomistic simulations
Zheyong Fan, Yanzhou Wang, Penghua Ying, Keke Song, Junjie Wang, Yong Wang, Zezhu Zeng, Ke Xu, Eric Lindgren, J Magnus Rahm, Alexander J Gabourie, Jiahui Liu, Haikuan Dong, Jianyang Wu, Yue Chen, Zheng Zhong, Jian Sun, Paul Erhart, Yanjing Su, Tapio Ala-Nissila
J. Chem. Phys., 2022, 157, 114801.
DOI: 10.1063/5.0106617
Magnetocaloric effect in ScGdTbDyHo high-entropy alloy: Impact of synthesis route
S.A. Uporov, S. Kh Estemirova, E.V. Sterkhov, I.A. Balyakin, A.A. Rempel
Intermetallics, 2022, 151, 107678.
DOI: 10.1016/j.intermet.2022.107678
A tungsten deep neural-network potential for simulating mechanical property degradation under fusion service environment
Xiaoyang Wang, Yinan Wang, Linfeng Zhang, Fuzhi Dai, Han Wang
Nucl. Fusion, 2022, 62, 126013.
DOI: 10.1088/1741-4326/ac888b
Molecular dynamics simulations of LiCl ion pairs in high temperature aqueous solutions by deep learning potential
Wei Zhang, Li Zhou, Bin Yang, Tinggui Yan
Journal of Molecular Liquids, 2022, 367, 120500.
DOI: 10.1016/j.molliq.2022.120500
DP Compress: A Model Compression Scheme for Generating Efficient Deep Potential Models
Denghui Lu, Wanrun Jiang, Yixiao Chen, Linfeng Zhang, Weile Jia, Han Wang, Mohan Chen
J. Chem. Theory Comput., 2022, 18, 5555–5567.
DOI: 10.1021/acs.jctc.2c00102
Combining Machine Learning Approaches and Accurate Ab Initio Enhanced Sampling Methods for Prebiotic Chemical Reactions in Solution
Timoth'ee Devergne, Th'eo Magrino, Fabio Pietrucci, A Marco Saitta
J. Chem. Theory Comput., 2022.
DOI: 10.1021/acs.jctc.2c00400
A Deep Neural Network Interface Potential for Li-Cu Systems
Genming Lai, Junyu Jiao, Chi Fang, Ruiqi Zhang, Xianqi Xu, Liyuan Sheng, Yao Jiang, Chuying Ouyang, Jiaxin Zheng
Adv. Mater. Interfaces, 2022, 2201346.
DOI: 10.1002/admi.202201346
Nucleation of Water Clusters in Gas Phase: A Computational Study Based on Neural Network Potential and Enhanced Sampling\textreferencemark
Sen Xu, Liling Wu, Zhenyu Li
Acta Chimica Sinica, 2022, 80, 598.
DOI: 10.6023/A22010003
Toward High-level Machine Learning Potential for Water Based on Quantum Fragmentation and Neural Networks
Jinfeng Liu, Jinggang Lan, Xiao He
J. Phys. Chem. A, 2022, 126, 3926–3936.
DOI: 10.1021/acs.jpca.2c00601
A Deep Neural Network Potential for Water Confined in Graphene Nanocapillaries
Wen Zhao, Hu Qiu, Wanlin Guo
J. Phys. Chem. C, 2022, 126, 10546–10553.
DOI: 10.1021/acs.jpcc.2c02423
Soft-phonon anharmonicity, floppy modes, and Na diffusion in Na3FY (Y=S,Se,Te): Ab initio and machine-learned molecular dynamics simulations
Mayanak Kumar Gupta, Sajan Kumar, Ranjan Mittal, Samrath L. Chaplot
Phys. Rev. B, 2022, 106, 14311.
DOI: 10.1103/PhysRevB.106.014311
Combined QM/MM, Machine Learning Path Integral Approach to Compute Free Energy Profiles and Kinetic Isotope Effects in RNA Cleavage Reactions
Timothy J Giese, Jinzhe Zeng, Şölen Ekesan, Darrin M York
J. Chem. Theory Comput., 2022, 18, 4304–4317.
DOI: 10.1021/acs.jctc.2c00151
Automated workflow for computation of redox potentials, acidity constants, and solvation free energies accelerated by machine learning
Feng Wang, Jun Cheng
J. Chem. Phys., 2022, 157, 24103.
DOI: 10.1063/5.0098330
Homogeneous ice nucleation in an ab initio machine-learning model of water
Pablo M Piaggi, Jack Weis, Athanassios Z Panagiotopoulos, Pablo G Debenedetti, Roberto Car
Proc. Natl. Acad. Sci. U. S. A., 2022, 119, e2207294119.
DOI: 10.1073/pnas.2207294119
High accuracy neural network interatomic potential for NiTi shape memory alloy
Hao Tang, Yin Zhang, Qing-Jie Li, Haowei Xu, Yuchi Wang, Yunzhi Wang, Ju Li
Acta Materialia, 2022, 238, 118217.
DOI: 10.1016/j.actamat.2022.118217
Machine Learning Accelerates Molten Salt Simulations: Thermal Conductivity of MgCl 2 -NaCl Eutectic
Wenshuo Liang, Guimin Lu, Jianguo Yu
Adv. Theory Simul., 2022, 2200206.
DOI: 10.1002/adts.202200206
Machine Learning Force Field Aided Cluster Expansion Approach to Configurationally Disordered Materials: Critical Assessment of Training Set Selection and Size Convergence
Jun-Zhong Xie, Xu-Yuan Zhou, Dong Luan, Hong Jiang
J. Chem. Theory Comput., 2022, 18, 3795–3804.
DOI: 10.1021/acs.jctc.2c00017
Combined Deep Learning and Classical Potential Approach for Modeling Diffusion in UiO-66
Siddarth K Achar, Jacob J Wardzala, Leonardo Bernasconi, Linfeng Zhang, J Karl Johnson
J. Chem. Theory Comput., 2022, 18, 3593–3606.
DOI: 10.1021/acs.jctc.2c00010
Deep neural network based quantum simulations and quasichemical theory for accurate modeling of molten salt thermodynamics
Yu Shi, Stephen T. Lam, Thomas~L. Beck
Chem. Sci., 2022.
DOI: 10.1039/D2SC02227C
Towards large-scale and spatiotemporally resolved diagnosis of electronic density of states by deep learning
Qiyu Zeng, Bo Chen, Xiaoxiang Yu, Shen Zhang, Dongdong Kang, Han Wang, Jiayu Dai
Phys. Rev. B, 2022, 105, 174109.
DOI: 10.1103/PhysRevB.105.174109
Exploring Complex Reaction Networks Using Neural Network-Based Molecular Dynamics Simulation
Qingzhao Chu, Kai H Luo, Dongping Chen
J. Phys. Chem. Lett., 2022, 13, 4052–4057.
DOI: 10.1021/acs.jpclett.2c00647
Reaction processes at step edges on S-decorated Cu(111) and Ag(111) surfaces: MD analysis utilizing machine learning derived potentials
Da-Jiang Liu, James W Evans
J. Chem. Phys., 2022, 156, 204106.
DOI: 10.1063/5.0089210
Deep machine learning potential for atomistic simulation of Fe-Si-O systems under Earth's outer core conditions
Chao Zhang, Ling Tang, Yang Sun, Kai-Ming Ho, Renata M. Wentzcovitch, Cai-Zhuang Wang
Phys. Rev. Materials, 2022, 6, 63802.
DOI: 10.1103/PhysRevMaterials.6.063802
Accelerated Deep Learning Dynamics for Atomic Layer Deposition of Al(Me)3 and Water on OH/Si(111)
Hiroya Nakata, Michael Filatov Gulak, Cheol Ho Choi
ACS Appl. Mater. Interfaces, 2022, 14, 26116–26127.
DOI: 10.1021/acsami.2c01768
Acids at the Edge: Why Nitric and Formic Acid Dissociations at Air-Water Interfaces Depend on Depth and on Interface Specific Area
Miguel de la Puente, Rolf David, Axel Gomez, Damien Laage
J. Am. Chem. Soc., 2022, 144, 10524–10529.
DOI: 10.1021/jacs.2c03099
Strongly Anharmonic Phonons and Their Role in Superionic Diffusion and Ultralow Thermal Conductivity of Cu 7 PSe 6
Mayanak K. Gupta, Jingxuan Ding, Dipanshu Bansal, Douglas L. Abernathy, Georg Ehlers, Naresh C. Osti, Wolfgang G. Zeier, Olivier Delaire
Advanced Energy Materials, 2022, 12, 2200596.
DOI: 10.1002/aenm.202200596
Atomistic Calculation of the Melting Point of the High-Entropy Cantor Alloy CoCrFeMnNi
I. A. Balyakin, A. A. Rempel
Dokl Phys Chem, 2022, 502, 11–17.
DOI: 10.1134/S0012501622010018
Deep potential development of transition-metal-rich carbides
Tyler McGilvry-James, Bikash Timalsina, Marium Mostafiz Mou, Ridwan Sakidja
MRS Advances, 2022, 7, 468–473.
DOI: 10.1557/s43580-022-00289-0
Accurate and efficient molecular dynamics based on machine learning and non von Neumann architecture
Pinghui Mo, Chang Li, Dan Zhao, Yujia Zhang, Mengchao Shi, Junhua Li, Jie Liu
npj Comput. Mater., 2022, 8, 107.
DOI: 10.1038/s41524-022-00773-z
Dissolving salt is not equivalent to applying a pressure on water
Chunyi Zhang, Shuwen Yue, Athanassios Z Panagiotopoulos, Michael L Klein, Xifan Wu
Nat. Commun., 2022, 13, 822.
DOI: 10.1038/s41467-022-28538-8
Exploring the Effects of Ionic Defects on the Stability of CsPbI 3 with a Deep Learning Potential
Weijie Yang, Jiajia Li, Xuelu Chen, Yajun Feng, Chongchong Wu, Ian D Gates, Zhengyang Gao, Xunlei Ding, Jianxi Yao, Hao Li
Chemphyschem, 2022, 23, e202100841.
DOI: 10.1002/cphc.202100841
Structural phase transitions in $\mathrmSrTi\mathrmO_3$ from deep potential molecular dynamics
Ri He, Hongyu Wu, Linfeng Zhang, Xiaoxu Wang, Fangjia Fu, Shi Liu, Zhicheng Zhong
Phys. Rev. B, 2022, 105, 064104.
DOI: 10.1103/PhysRevB.105.064104

Efficient and accurate atomistic modeling of dopant migration using deep neural network
Xi Ding, Ming Tao, Junhua Li, Mingyuan Li, Mengchao Shi, Jiashu Chen, Zhen Tang, Francis Benistant, Jie Liu
Materials Science in Semiconductor Processing, 2022, 143, 106513.
DOI: 10.1016/j.mssp.2022.106513

Self-Healing Mechanism of Lithium in Lithium Metal
Junyu Jiao, Genming Lai, Liang Zhao, Jiaze Lu, Qidong Li, Xianqi Xu, Yao Jiang, Yan-Bing He, Chuying Ouyang, Feng Pan, Hong Li, Jiaxin Zheng
Adv. Sci. (Weinh)., 2022, 9, e2105574.
DOI: 10.1002/advs.202105574

A deep learning interatomic potential developed for atomistic simulation of carbon materials
Jinjin Wang, Hong Shen, Riyi Yang, Kun Xie, Chao Zhang, Liangyao Chen, Kai-Ming Ho, Cai-Zhuang Wang, Songyou Wang
Carbon, 2022, 186, 1–8.
DOI: 10.1016/j.carbon.2021.09.062

Using metadynamics to build neural network potentials for reactive events: the case of urea decomposition in water
Manyi Yang, Luigi Bonati, Daniela Polino, Michele Parrinello
Catalysis Today, 2022, 387, 143–149.
DOI: 10.1016/j.cattod.2021.03.018

Development of neural network potential for MD simulation and its application to TiN
Takeru Miyagawa, Kazuki Mori, Nobuhiko Kato, Akio Yonezu
Computational Materials Science, 2022, 206, 111303.
DOI: 10.1016/j.commatsci.2022.111303

Ab Initio Neural Network MD Simulation of Thermal Decomposition of High Energy Material CL-20/TNT
Liqun Cao, Jinzhe Zeng, Bo Wang, Tong Zhu, John Z.H. Zhang
Physical Chemistry Chemical Physics, 2022, 24, 11801–11811.
DOI: 10.1039/D2CP00710J
Temperature- and vacancy-concentration-dependence of heat transport in Li3ClO from multi-method numerical simulations
Paolo Pegolo, Stefano Baroni, Federico Grasselli
npj Comput. Mater., 2022, 8, 24.
DOI: 10.1038/s41524-021-00693-4
Grain boundary segregation induced strong UHTCs at elevated temperatures: A universal mechanism from conventional UHTCs to high entropy UHTCs
Fu-Zhi Dai, Bo Wen, Yinjie Sun, Yixiao Ren, Huimin Xiang, Yanchun Zhou
Journal of Materials Science & Technology, 2022, 123, 26-33.
DOI: 10.1016/j.jmst.2021.12.074
The chemical origin of temperature-dependent lithium-ion concerted diffusion in sulfide solid electrolyte Li10GeP2S12
Zhong-Heng Fu, Xiang Chen, Nan Yao, Xin Shen, Xia-Xia Ma, Shuai Feng, Shuhao Wang, Rui Zhang, Linfeng Zhang, Qiang Zhang
Journal of Energy Chemistry, 2022, 70, 59–66.
DOI: 10.1016/j.jechem.2022.01.018
Study on the structural properties of refining slags by molecular dynamics with deep learning potential
Yuhan Sun, Min Tan, Tao Li, Junguo Li, Bo Shang
Journal of Molecular Liquids, 2022, 353, 118787.
DOI: 10.1016/j.molliq.2022.118787
Nanotwinning induced decreased lattice thermal conductivity of high temperature thermoelectric boron subphosphide (B12P2) from deep learning potential simulations
Xiaona Huang, Yidi Shen, Qi An
Energy and AI, 2022, 8, 100135.
DOI: 10.1016/j.egyai.2022.100135
A deep potential model with long-range electrostatic interactions
Linfeng Zhang, Han Wang, Maria Carolina Muniz, Athanassios Z Panagiotopoulos, Roberto Car, Weinan E
J. Chem. Phys., 2022, 156, 124107.
DOI: 10.1063/5.0083669
Four-Phonon Scattering Effect and Two-Channel Thermal Transport in Two-Dimensional Paraelectric SnSe
Jie Sun, Cunzhi Zhang, Zhonghua Yang, Yiheng Shen, Ming Hu, Qian Wang
ACS Appl. Mater. Interfaces, 2022, 14, 11493–11499.
DOI: 10.1021/acsami.1c24488
An inductive transfer learning force field (ITLFF) protocol builds protein force fields in seconds
Yanqiang Han, Zhilong Wang, An Chen, Imran Ali, Junfei Cai, Simin Ye, Jinjin Li
Brief. Bioinform., 2022, 23.
DOI: 10.1093/bib/bbab590
Sulfur-enhanced dynamics of coinage metal(111) surfaces: Step edges versus terraces as locations for metal-sulfur complex formation
Da-Jiang Liu, James W. Evans
Journal of Vacuum Science \& Technology A, 2022, 40 (2), 023205.
DOI: 10.1116/6.0001408
A generalizable machine learning potential of Ag-Au nanoalloys and its application to surface reconstruction, segregation and diffusion
YiNan Wang, LinFeng Zhang, Ben Xu, XiaoYang Wang, Han Wang
Modelling Simul. Mater. Sci. Eng., 2022, 30, 25003.
DOI: 10.1088/1361-651X/ac4002
Deep machine learning potentials for multicomponent metallic melts: Development, predictability and compositional transferability
R.E. Ryltsev, N.M. Chtchelkatchev
Journal of Molecular Liquids, 2022, 349, 118181.
DOI: 10.1016/j.molliq.2021.118181
Neural network potential for Zr-Rh system by machine learning
Kun Xie, Chong Qiao, Hong Shen, Riyi Yang, Ming Xu, Chao Zhang, Yuxiang Zheng, Rongjun Zhang, Liangyao Chen, Kai-Ming Ho, Cai-Zhuang Wang, Songyou Wang
J. Phys. Condens. Matter, 2022, 34, 75402.
DOI: 10.1088/1361-648X/ac37dc
Molecular dynamics simulation of molten strontium chloride based on deep potential
Di Guo, Jia Zhao, Wenshuo Liang, Guimin Lu
Journal of Molecular Liquids, 2022, 348, 118380.
DOI: 10.1016/j.molliq.2021.118380
Suppression of Rayleigh Scattering in Silica Glass by Codoping Boron and Fluorine: Molecular Dynamics Simulations with Force-Matching and Neural Network Potentials
Shingo Urata, Nobuhiro Nakamura, Tomofumi Tada, Aik Rui Tan, Rafael Gómez-Bombarelli, Hideo Hosono
J. Phys. Chem. C, 2022, 126 (4), 2264-2275.
DOI: 10.1021/acs.jpcc.1c10300
A deep learning potential applied in tobermorite phases and extended to calcium silicate hydrates
Yang Zhou, Haojie Zheng, Weihuan Li, Tao Ma, Changwen Miao
Cement and Concrete Research, 2022, 152, 106685.
DOI: 10.1016/j.cemconres.2021.106685
Deep learning potential for superionic phase of Ag2S
I.A. Balyakin, S.I. Sadovnikov
Computational Materials Science, 2022, 202, 110963.
DOI: 10.1016/j.commatsci.2021.110963
Neural network representation of electronic structure from ab initio molecular dynamics
Qiangqiang Gu, Linfeng Zhang, Ji Feng
Science Bulletin, 2022, 67, 29–37.
DOI: 10.1016/j.scib.2021.09.010
2021
Machine learning builds full-QM precision protein force fields in seconds
Yanqiang Han, Zhilong Wang, Zhiyun Wei, Jinyun Liu, Jinjin Li
Brief. Bioinform., 2021, 22.
DOI: 10.1093/bib/bbab158

Efficiently Trained Deep Learning Potential for Graphane
Siddarth K. Achar, Linfeng Zhang, J. Karl Johnson
J. Phys. Chem. C, 2021, 125, 14874–14882.
DOI: 10.1021/acs.jpcc.1c01411

2D Heterostructure of Amorphous CoFeB Coating Black Phosphorus Nanosheets with Optimal Oxygen Intermediate Absorption for Improved Electrocatalytic Water Oxidation
Huayu Chen, Junxiang Chen, Pei Ning, Xin Chen, Junhui Liang, Xin Yao, Da Chen, Laishun Qin, Yuexiang Huang, Zhenhai Wen
ACS Nano, 2021, 15, 12418–12428.
DOI: 10.1021/acsnano.1c04715

Deep potential generation scheme and simulation protocol for the Li10GeP2S12-type superionic conductors
Jianxing Huang, Linfeng Zhang, Han Wang, Jinbao Zhao, Jun Cheng, Weinan E
J. Chem. Phys., 2021, 154, 94703.
DOI: 10.1063/5.0041849

Machine-Learning-Assisted Free Energy Simulation of Solution-Phase and Enzyme Reactions
Xiaoliang Pan, Junjie Yang, Richard Van, Evgeny Epifanovsky, Junming Ho, Jing Huang, Jingzhi Pu, Ye Mei, Kwangho Nam, Yihan Shao
J. Chem. Theory Comput., 2021, 17, 5745–5758.
DOI: 10.1021/acs.jctc.1c00565
Accurate force field of two-dimensional ferroelectrics from deep learning
Jing Wu, Liyi Bai, Jiawei Huang, Liyang Ma, Jian Liu, Shi Liu
Phys. Rev. B, 2021, 104, 174107.
DOI: 10.1103/PhysRevB.104.174107

Exploring the Chemical Space of Linear Alkane Pyrolysis via Deep Potential GENerator
Jinzhe Zeng, Linfeng Zhang, Han Wang, Tong Zhu
Energy & Fuels, 2021, 35 (1), 762–769.
DOI: 10.1021/acs.energyfuels.0c03211
Development of Range-Corrected Deep Learning Potentials for Fast, Accurate Quantum Mechanical/Molecular Mechanical Simulations of Chemical Reactions in Solution
Jinzhe Zeng, Timothy J Giese, Şölen Ekesan, Darrin M York
J. Chem. Theory Comput., 2021, 17, 6993–7009.
DOI: 10.1021/acs.jctc.1c00201

Reactive uptake of N2O5 by atmospheric aerosol is dominated by interfacial processes
Mirza Galib, David T Limmer
Science, 2021, 371, 921–925.
DOI: 10.1126/science.abd7716

86 PFLOPS Deep Potential Molecular Dynamics simulation of 100 million atoms with ab initio accuracy
Denghui Lu, Han Wang, Mohan Chen, Lin Lin, Roberto Car, Weinan E, Weile Jia, Linfeng Zhang
Computer Physics Communications, 2021, 259, 107624.
DOI: 10.1016/j.cpc.2020.107624

Insights from Computational Studies on the Anisotropic Volume Change of LixNiO2 at High States of Charge (x < 0.25)
Juan C. Garcia, Joshua Gabriel, Noah H. Paulson, John Low, Marius Stan, Hakim Iddir
J. Phys. Chem. C, 2021, 125 (49), 27130-27139.
DOI: 10.1021/acs.jpcc.1c08022

Thermodynamic and Transport Properties of LiF and FLiBe Molten Salts with Deep Learning Potentials
Alejandro Rodriguez, Stephen Lam, Ming Hu
ACS Appl. Mater. Interfaces, 2021, 13, 55367–55379.
DOI: 10.1021/acsami.1c17942
Heat transport in liquid water from first-principles and deep neural network simulations
Davide Tisi, Linfeng Zhang, Riccardo Bertossa, Han Wang, Roberto Car, Stefano Baroni
Phys. Rev. B, 2021, 104, 224202.
DOI: 10.1103/PhysRevB.104.224202
Specialising neural network potentials for accurate properties and application to the mechanical response of titanium
Tongqi Wen, Rui Wang, Lingyu Zhu, Linfeng Zhang, Han Wang, David J. Srolovitz, Zhaoxuan Wu
npj Comput. Mater., 2021, 7, 206.
DOI: 10.1038/s41524-021-00661-y
Fast Na diffusion and anharmonic phonon dynamics in superionic Na3PS4
Mayanak K. Gupta, Jingxuan Ding, Naresh C. Osti, Douglas L. Abernathy, William Arnold, Hui Wang, Zachary Hood, Olivier Delaire
Energy Environ. Sci., 2021, 14, 6554-6563.
DOI: 10.1039/D1EE01509E
Experimental observation of localized interfacial phonon modes
Zhe Cheng, Ruiyang Li, Xingxu Yan, Glenn Jernigan, Jingjing Shi, Michael E Liao, Nicholas J Hines, Chaitanya A Gadre, Juan Carlos Idrobo, Eungkyu Lee, Karl D Hobart, Mark S Goorsky, Xiaoqing Pan, Tengfei Luo, Samuel Graham
Nat. Commun., 2021, 12, 6901.
DOI: 10.1038/s41467-021-27250-3

Artificial intelligence model for efficient simulation of monatomic phase change material antimony
Mengchao Shi, Junhua Li, Ming Tao, Xin Zhang, Jie Liu
Materials Science in Semiconductor Processing, 2021, 136, 106146.
DOI: 10.1016/j.mssp.2021.106146

Molecular dynamics simulation of metallic Al-Ce liquids using a neural network machine learning interatomic potential
L Tang, K M Ho, C Z Wang
J. Chem. Phys., 2021, 155, 194503.
DOI: 10.1063/5.0066061
Choosing the right molecular machine learning potential
Max Pinheiro Jr, Fuchun Ge, Nicolas Ferr'e, Pavlo O Dral, Mario Barbatti
Chem. Sci., 2021, 12, 14396–14413.
DOI: 10.1039/d1sc03564a

Atomic structure of liquid refractory Nb5Si3 intermetallic compound alloy based upon deep neural network potential
Q. Wang, B. Zhai, H. P. Wang, B. Wei
Journal of Applied Physics, 2021, 130, 185103.
DOI: 10.1063/5.0067157

Azo(xy) vs Aniline Selectivity in Catalytic Nitroarene Reduction by Intermetallics: Experiments and Simulations
Carena L. Daniels, Da-Jiang Liu, Marquix A. S. Adamson, Megan Knobeloch, Javier Vela
J. Phys. Chem. C, 2021, 125 (44), 24440-24450.
DOI: 10.1021/acs.jpcc.1c08569

Resolving the Structural Debate for the Hydrated Excess Proton in Water
Paul B Calio, Chenghan Li, Gregory A Voth
J. Am. Chem. Soc., 2021, 143, 18672–18683.
DOI: 10.1021/jacs.1c08552
Deep-learning potential method to simulate shear viscosity of liquid aluminum at high temperature and high pressure by molecular dynamics
Yuqing Cheng, Han Wang, Shuaichuang Wang, Xingyu Gao, Qiong Li, Jun Fang, Hongzhou Song, Weidong Chu, Gongmu Zhang, Haifeng Song, Haifeng Liu
AIP Advances, 2021, 11, 15043.
DOI: 10.1063/5.0036298
Gold Segregation Improves Electrocatalytic Activity of Icosahedron Au@Pt Nanocluster: Insights from Machine Learning
Dingming Chen, Zhuangzhuang Lai, Jiawei Zhang, Jianfu Chen, Peijun Hu, Haifeng Wang
Chin. J. Chem., 2021, 39, 3029–3036.
DOI: 10.1002/cjoc.202100352
Condensed Phase Water Molecular Multipole Moments from Deep Neural Network Models Trained on Ab Initio Simulation Data
Yu Shi, Carrie C Doyle, Thomas L Beck
J. Phys. Chem. Lett., 2021, 12, 10310–10317.
DOI: 10.1021/acs.jpclett.1c02328
Learning intermolecular forces at liquid-vapor interfaces
Samuel P Niblett, Mirza Galib, David T Limmer
J. Chem. Phys., 2021, 155, 164101.
DOI: 10.1063/5.0067565
Modeling Liquid Water by Climbing up Jacob’s Ladder in Density Functional Theory Facilitated by Using Deep Neural Network Potentials
Chunyi Zhang, Fujie Tang, Mohan Chen, Jianhang Xu, Linfeng Zhang, Diana Y Qiu, John P Perdew, Michael L Klein, Xifan Wu
J. Phys. Chem. B, 2021, 125, 11444–11456.
DOI: 10.1021/acs.jpcb.1c03884
Deep Density: Circumventing the Kohn-Sham equations via symmetry preserving neural networks
Leonardo Zepeda-N'u\~nez, Yixiao Chen, Jiefu Zhang, Weile Jia, Linfeng Zhang, Lin Lin
Journal of Computational Physics, 2021, 443, 110523.
DOI: 10.1016/j.jcp.2021.110523
First-principles materials simulation and design for alkali and alkaline metal ion batteries accelerated by machine learning
Lujie Jin, Yujin Ji, Hongshuai Wang, Lifeng Ding, Youyong Li
Phys. Chem. Chem. Phys., 2021, 23, 21470–21483.
DOI: 10.1039/d1cp02963k
Local structure elucidation and properties prediction on KCl-CaCl2 molten salt: A deep potential molecular dynamics study
Min Bu, Wenshuo Liang, Guimin Lu, Jianguo Yu
Solar Energy Materials and Solar Cells, 2021, 232, 111346.
DOI: 10.1016/j.solmat.2021.111346
Using Neural Network Force Fields to Ascertain the Quality of Ab Initio Simulations of Liquid Water
Alberto Torres, Luana S Pedroza, Marivi Fernandez-Serra, Alexandre R Rocha
J. Phys. Chem. B, 2021, 125, 10772–10778.
DOI: 10.1021/acs.jpcb.1c04372
Thermal Conductivity of Silicate Liquid Determined by Machine Learning Potentials
Jie Deng, Lars Stixrude
Geophys Res Lett, 2021, 48, e2021GL093806.
DOI: 10.1029/2021GL093806
Ab initio validation on the connection between atomistic and hydrodynamic description to unravel the ion dynamics of warm dense matter
Qiyu Zeng, Xiaoxiang Yu, Yunpeng Yao, Tianyu Gao, Bo Chen, Shen Zhang, Dongdong Kang, Han Wang, Jiayu Dai
Phys. Rev. Research, 2021, 3, 33116.
DOI: 10.1103/PhysRevResearch.3.033116
Liquid-Liquid Critical Point in Phosphorus
Manyi Yang, Tarak Karmakar, Michele Parrinello
Phys. Rev. Lett., 2021, 127, 80603.
DOI: 10.1103/PhysRevLett.127.080603
Robust, Multi-Length-Scale, Machine Learning Potential for Ag–Au Bimetallic Alloys from Clusters to Bulk Materials
Christopher M. Andolina, Marta Bon, Daniele Passerone, Wissam A. Saidi
J. Phys. Chem. C, 2021, 125 (31), 17438-17447.
DOI: 10.1021/acs.jpcc.1c04403
Improved Al-Mg alloy surface segregation predictions with a machine learning atomistic potential
Md Sabbir Akhanda, S Emad Rezaei, Keivan Esfarjani, Sergiy Krylyuk, Albert V Davydov, Mona Zebarjadi
Phys. Rev. Mater., 2021, 5, 83804.
DOI: 10.1103/PhysRevMaterials.5.083804
Anomalous Behavior of Viscosity and Electrical Conductivity of MgSiO 3 Melt at Mantle Conditions
Haiyang Luo, Bijaya B. Karki, Dipta B. Ghosh, Huiming Bao
Geophys Res Lett, 2021, 48.
DOI: 10.1029/2021GL093573
Deep neural network potentials for diffusional lithium isotope fractionation in silicate melts
Haiyang Luo, Bijaya B. Karki, Dipta B. Ghosh, Huiming Bao
Geochimica et Cosmochimica Acta, 2021, 303, 38–50.
DOI: 10.1016/j.gca.2021.03.031
Phase Diagram of a Deep Potential Water Model
Linfeng Zhang, Han Wang, Roberto Car, Weinan E
Phys. Rev. Lett., 2021, 126, 236001.
DOI: 10.1103/PhysRevLett.126.236001
Theoretical prediction on the local structure and transport properties of molten alkali chlorides by deep potentials
Wenshuo Liang, Guimin Lu, Jianguo Yu
Journal of Materials Science & Technology, 2021, 75, 78-85.
DOI: 10.1016/j.jmst.2020.09.040
The thermoelectric performance of new structure SnSe studied by quotient graph and deep learning potential
D. Guo, C. Li, K. Li, B. Shao, D. Chen, Y. Ma, J. Sun, X. Cao, W. Zeng, X. Chang
Materials Today Energy, 2021, 20, 100665.
DOI: 10.1016/j.mtener.2021.100665
Phase Equilibrium of Water with Hexagonal and Cubic Ice Using the SCAN Functional
Pablo M Piaggi, Athanassios Z Panagiotopoulos, Pablo G Debenedetti, Roberto Car
J. Chem. Theory Comput., 2021, 17, 3065–3077.
DOI: 10.1021/acs.jctc.1c00041
Temperature Dependent Thermal and Elastic Properties of High Entropy (Ti0.2Zr0.2Hf0.2Nb0.2Ta0.2)B2: Molecular Dynamics Simulation by Deep Learning Potential
Fu-Zhi Dai, Yinjie Sun, Bo Wen, Huimin Xiang, Yanchun Zhou
Journal of Materials Science & Technology, 2021, 72, 8-15.
DOI: 10.1016/j.jmst.2020.07.014
Theoretical prediction on the redox potentials of rare-earth ions by deep potentials
Jia Zhao, Wenshuo Liang, Guimin Lu
Ionics, 2021, 27, 2079–2088.
DOI: 10.1007/s11581-021-03988-0
Accurate Deep Potential model for the Al-Cu-Mg alloy in the full concentration space*
Wanrun Jiang, Yuzhi Zhang, Linfeng Zhang, Han Wang
Chinese Phys. B, 2021, 30, 50706.
DOI: 10.1088/1674-1056/abf134
Anharmonic Raman spectra simulation of crystals from deep neural networks
Honghui Shang, Haidi Wang
AIP Advances, 2021, 11, 35105.
DOI: 10.1063/5.0040190
Thermal transport by electrons and ions in warm dense aluminum: A combined density functional theory and deep potential study
Qianrui Liu, Junyi Li, Mohan Chen
Matter and Radiation at Extremes, 2021, 6 (2), 026902.
DOI: 10.1063/5.0030123
Crystallization of the P3Sn4 Phase upon Cooling P2Sn5 Liquid by Molecular Dynamics Simulation Using a Machine Learning Interatomic Potential
Chao Zhang, Yang Sun, Hai-Di Wang, Feng Zhang, Tong-Qi Wen, Kai-Ming Ho, Cai-Zhuang Wang
J. Phys. Chem. C, 2021, 125 (5), 3127-3133.
DOI: 10.1021/acs.jpcc.0c08873
Enhancing the formation of ionic defects to study the ice Ih/XI transition with molecular dynamics simulations
Pablo M. Piaggi, Roberto Car
Molecular Physics, 2021, 119.
DOI: 10.1080/00268976.2021.1916634
Static and Dynamic Correlations in Water: Comparison of Classical Ab Initio Molecular Dynamics at Elevated Temperature with Path Integral Simulations at Ambient Temperature
Chenghan Li, Francesco Paesani, Gregory A Voth
J. Chem. Theory Comput., 2022, 18, 2124–2131.
DOI: 10.1021/acs.jctc.1c01223
Molecular dynamics simulations of lanthanum chloride by deep learning potential
Taixi Feng, Jia Zhao, Wenshuo Liang, Guimin Lu
Computational Materials Science, 2021, 111014.
DOI: 10.1016/j.commatsci.2021.111014
Diffusional fractionation of helium isotopes in silicate melts
H. Luo, B.B. Karki, D.B. Ghosh, H. Bao
Geochem. Persp. Let., 2021, 19–22.
DOI: 10.7185/geochemlet.2128
Short- and medium-range orders in Al90Tb10 glass and their relation to the structures of competing crystalline phases
L. Tang, Z.J. Yang, T.Q. Wen, K.M. Ho, M.J. Kramer, C.Z. Wang
Acta Materialia, 2021, 204, 116513.
DOI: 10.1016/j.actamat.2020.116513
A DFT accurate machine learning description of molten ZnCl2 and its mixtures: 2. Potential development and properties prediction of ZnCl2-NaCl-KCl ternary salt for CSP
Gechuanqi Pan, Jing Ding, Yunfei Du, Duu-Jong Lee, Yutong Lu
Computational Materials Science, 2021, 187, 110055.
DOI: 10.1016/j.commatsci.2020.110055
Deep learning of accurate force field of ferroelectric HfO2
Jing Wu, Yuzhi Zhang, Linfeng Zhang, Shi Liu
Phys. Rev. B, 2021, 103, 24108.
DOI: 10.1103/PhysRevB.103.024108
Machine-Learning-Driven Simulations on Microstructure and Thermophysical Properties of MgCl2-KCl Eutectic
Wenshuo Liang, Guimin Lu, Jianguo Yu
ACS Appl. Mater. Interfaces, 2021, 13, 4034–4042.
DOI: 10.1021/acsami.0c20665

Theoretical study of Na+ transport in the solid-state electrolyte Na3OBr based on deep potential molecular dynamics
Han-Xiao Li, Xu-Yuan Zhou, Yue-Chao Wang, Hong Jiang
Inorg. Chem. Front., 2021, 8, 425–432.
DOI: 10.1039/D0QI00921K
When do short-range atomistic machine-learning models fall short?
Shuwen Yue, Maria Carolina Muniz, Marcos F Calegari Andrade, Linfeng Zhang, Roberto Car, Athanassios Z Panagiotopoulos
J. Chem. Phys., 2021, 154, 34111.
DOI: 10.1063/5.0031215
2020
Simulating Diffusion Properties of Solid-State Electrolytes via a Neural Network Potential: Performance and Training Scheme
Aris Marcolongo, Tobias Binninger, Federico Zipoli, Teodoro Laino
Chemsystemschem, 2020, 2 (3).
DOI: 10.1002/syst.201900031
Free energy of proton transfer at the water-TiO2 interface from ab initio deep potential molecular dynamics
Marcos F Calegari Andrade, Hsin-Yu Ko, Linfeng Zhang, Roberto Car, Annabella Selloni
Chem. Sci., 2020, 11, 2335–2341.
DOI: 10.1039/C9SC05116C

Theoretical prediction on thermal and mechanical properties of high entropy (Zr0.2Hf0.2Ti0.2Nb0.2Ta0.2)C by deep learning potential
Fu-Zhi Dai, Bo Wen, Yinjie Sun, Huimin Xiang, Yanchun Zhou
Journal of Materials Science & Technology, 2020, 43, 168–174.
DOI: 10.1016/j.jmst.2020.01.005

A deep neural network interatomic potential for studying thermal conductivity of $\beta$-Ga2O3
Ruiyang Li, Zeyu Liu, Andrew Rohskopf, Kiarash Gordiz, Asegun Henry, Eungkyu Lee, Tengfei Luo
Appl. Phys. Lett., 2020, 117, 152102.
DOI: 10.1063/5.0025051

Structure and dynamics of warm dense aluminum: a molecular dynamics study with density functional theory and deep potential
Qianrui Liu, Denghui Lu, Mohan Chen
J. Phys. Condens. Matter, 2020, 32, 144002.
DOI: 10.1088/1361-648X/ab5890
Ab initio phase diagram and nucleation of gallium
Haiyang Niu, Luigi Bonati, Pablo M Piaggi, Michele Parrinello
Nat. Commun., 2020, 11, 2654.
DOI: 10.1038/s41467-020-16372-9
Raman spectrum and polarizability of liquid water from deep neural networks
Grace M Sommers, Marcos F Calegari Andrade, Linfeng Zhang, Han Wang, Roberto Car
Phys. Chem. Chem. Phys., 2020, 22, 10592–10602.
DOI: 10.1039/D0CP01893G

A machine learning based deep potential for seeking the low-lying candidates of Al clusters
P Tuo, X B Ye, B C Pan
J. Chem. Phys., 2020, 152, 114105.
DOI: 10.1063/5.0001491
Data-driven coarse-grained modeling of polymers in solution with structural and dynamic properties conserved
Shu Wang, Zhan Ma, Wenxiao Pan
Soft Matter, 2020, 16, 8330–8344.
DOI: 10.1039/D0SM01019G

Complex reaction processes in combustion unraveled by neural network-based molecular dynamics simulation
Jinzhe Zeng, Liqun Cao, Mingyuan Xu, Tong Zhu, John Z. H. Zhang
Nature Communications, 2020, 11, 5713.
DOI: 10.1038/s41467-020-19497-z
Deep neural network for the dielectric response of insulators
Linfeng Zhang, Mohan Chen, Xifan Wu, Han Wang, Weinan E, Roberto Car
Phys. Rev. B, 2020, 102, 41121.
DOI: 10.1103/PhysRevB.102.041121

DP-GEN: A concurrent learning platform for the generation of reliable deep learning based potential energy models
Yuzhi Zhang, Haidi Wang, Weijie Chen, Jinzhe Zeng, Linfeng Zhang, Han Wang, Weinan E
Computer Physics Communications, 2020, 253, 107206.
DOI: 10.1016/j.cpc.2020.107206
Isotope effects in molecular structures and electronic properties of liquid water via deep potential molecular dynamics based on the SCAN functional
Jianhang Xu, Chunyi Zhang, Linfeng Zhang, Mohan Chen, Biswajit Santra, Xifan Wu
Phys. Rev. B, 2020, 102, 214113.
DOI: 10.1103/PhysRevB.102.214113

Hydrogen Dynamics in Supercritical Water Probed by Neutron Scattering and Computer Simulations
Carla Andreani, Giovanni Romanelli, Alexandra Parmentier, Roberto Senesi, Alexander I Kolesnikov, Hsin-Yu Ko, Marcos F Calegari Andrade, Roberto Car
J. Phys. Chem. Lett., 2020, 11, 9461–9467.
DOI: 10.1021/acs.jpclett.0c02547

A unified deep neural network potential capable of predicting thermal conductivity of silicon in different phases
R. Li, E. Lee, T. Luo
Materials Today Physics, 2020, 12, 100181.
DOI: 10.1016/j.mtphys.2020.100181

Combining the Fragmentation Approach and Neural Network Potential Energy Surfaces of Fragments for Accurate Calculation of Protein Energy
Zhilong Wang, Yanqiang Han, Jinjin Li, Xiao He
J. Phys. Chem. B, 2020, 124, 3027–3035.
DOI: 10.1021/acs.jpcb.0c01370

A Deep-Learning Potential for Crystalline and Amorphous Li-Si Alloys
Nan Xu, Yao Shi, Yi He, Qing Shao
J. Phys. Chem. C, 2020, 124, 16278–16288.
DOI: 10.1021/acs.jpcc.0c03333
Development of interatomic potential for Al-Tb alloys using a deep neural network learning method
L Tang, Z J Yang, T Q Wen, K M Ho, M J Kramer, C Z Wang
Phys. Chem. Chem. Phys., 2020, 22, 18467–18479.
DOI: 10.1039/D0CP01689F
Isotope effects in x-ray absorption spectra of liquid water
Chunyi Zhang, Linfeng Zhang, Jianhang Xu, Fujie Tang, Biswajit Santra, Xifan Wu
Phys. Rev. B, 2020, 102, 115155.
DOI: 10.1103/PhysRevB.102.115155

Warm dense matter simulation via electron temperature dependent deep potential molecular dynamics
Yuzhi Zhang, Chang Gao, Qianrui Liu, Linfeng Zhang, Han Wang, Mohan Chen
Physics of Plasmas, 2020, 27, 122704.
DOI: 10.1063/5.0023265

Molecular Dynamics Simulations of Molten Magnesium Chloride Using Machine-Learning-Based Deep Potential
Wenshuo Liang, Guimin Lu, Jianguo Yu
Adv. Theory Simul., 2020, 3, 2000180.
DOI: 10.1002/adts.202000180

Grain boundary strengthening in ZrB2 by segregation of W: Atomistic simulations with deep learning potential
Fu-Zhi Dai, Bo Wen, Huimin Xiang, Yanchun Zhou
Journal of the European Ceramic Society, 2020, 40, 5029–5036.
DOI: 10.1016/j.jeurceramsoc.2020.06.007
Crystal Structure Prediction of Binary Alloys via Deep Potential
Haidi Wang, Yuzhi Zhang, Linfeng Zhang, Han Wang
Front. Chem., 2020, 8, 589795.
DOI: 10.3389/fchem.2020.589795
Deep machine learning interatomic potential for liquid silica
I A Balyakin, S V Rempel, R E Ryltsev, A A Rempel
Phys. Rev. E, 2020, 102, 52125.
DOI: 10.1103/PhysRevE.102.052125
Structure of disordered TiO2 phases from ab initio based deep neural network simulations
Marcos F. Calegari Andrade, Annabella Selloni
Phys. Rev. Materials, 2020, 4, 113803.
DOI: 10.1103/PhysRevMaterials.4.113803

Signatures of a liquid-liquid transition in an ab initio deep neural network model for water
Thomas E Gartner 3rd, Linfeng Zhang, Pablo M Piaggi, Roberto Car, Athanassios Z Panagiotopoulos, Pablo G Debenedetti
Proc. Natl. Acad. Sci. U. S. A., 2020, 117, 26040–26046.
DOI: 10.1073/pnas.2015440117
2019
Active learning of uniformly accurate interatomic potentials for materials simulation
Linfeng Zhang, De-Ye Lin, Han Wang, Roberto Car, Weinan E
Phys. Rev. Materials, 2019, 3, 23804.
DOI: 10.1103/PhysRevMaterials.3.023804

Isotope effects in liquid water via deep potential molecular dynamics
Hsin-Yu Ko, Linfeng Zhang, Biswajit Santra, Han Wang, Weinan E, Robert A. DiStasio Jr, Roberto Car
Molecular Physics, 2019, 117, 3269–3281.
DOI: 10.1080/00268976.2019.1652366
Development of a deep machine learning interatomic potential for metalloid-containing Pd-Si compounds
Tongqi Wen, Cai-Zhuang Wang, M. J. Kramer, Yang Sun, Beilin Ye, Haidi Wang, Xueyuan Liu, Chao Zhang, Feng Zhang, Kai-Ming Ho, Nan Wang
Phys. Rev. B, 2019, 100, 174101.
DOI: 10.1103/PhysRevB.100.174101

Deep learning inter-atomic potential model for accurate irradiation damage simulations
Hao Wang, Xun Guo, Linfeng Zhang, Han Wang, Jianming Xue
Appl. Phys. Lett., 2019, 114, 244101.
DOI: 10.1063/1.5098061
2018
Silicon Liquid Structure and Crystal Nucleation from Ab~Initio Deep Metadynamics
Luigi Bonati, Michele Parrinello
Phys. Rev. Lett., 2018, 121, 265701.
DOI: 10.1103/PhysRevLett.121.265701

Deep Learning for Nonadiabatic Excited-State Dynamics
Wen-Kai Chen, Xiang-Yang Liu, Wei-Hai Fang, Pavlo O Dral, Ganglong Cui
J. Phys. Chem. Lett., 2018, 9, 6702–6708.
DOI: 10.1021/acs.jpclett.8b03026

Adaptive coupling of a deep neural network potential to a classical force field
Linfeng Zhang, Han Wang, Weinan E
J. Chem. Phys., 2018, 149, 154107.
DOI: 10.1063/1.5042714

DeePCG: Constructing coarse-grained models via deep neural networks
Linfeng Zhang, Jiequn Han, Han Wang, Roberto Car, Weinan E
J. Chem. Phys., 2018, 149, 34101.
DOI: 10.1063/1.5027645

DeePMD-kit: A deep learning package for many-body potential energy representation and molecular dynamics
Han Wang, Linfeng Zhang, Jiequn Han, Weinan E
Computer Physics Communications, 2018, 228, 178–184.
DOI: 10.1016/j.cpc.2018.03.016
Deep Potential Molecular Dynamics: A Scalable Model with the Accuracy of Quantum Mechanics
Linfeng Zhang, Jiequn Han, Han Wang, Roberto Car, Weinan E
Phys. Rev. Lett., 2018, 120, 143001.
DOI: 10.1103/PhysRevLett.120.143001